Related Experiment Video
Updated: Jan 15, 2026

Electrocardiogram Recordings in Anesthetized Mice using Lead II
Published on: June 20, 2020
Assessment of QT Interval Prolongation Using Concentration-QT Modeling for Iptacopan, an Oral Complement Factor B
Noemi Kaetterer1, Kenneth Kulmatycki2, Prasanna Kumar Nidamarthy3
1BioMedical Research, Novartis Pharma AG, Basel, Switzerland.

To assess cardiac and safety parameters of iptacopan (an oral, selective, reversible, small-molecule factor B inhibitor), we conducted a phase I, single ascending dose (SAD), exposure-response study (A2107) instead of a traditional thorough QT study. Healthy participants were randomized 3:1 to receive a single, supratherapeutic, oral dose of iptacopan 400, 800, or 1200 mg, or placebo. A2107's primary objectives were to assess iptacopan's effect on Fridericia-corrected QT interval (QTcF) in a pooled analysis with the SAD phase of the first-in-human X2101 study, and the safety and tolerability of supratherapeutic iptacopan doses in participants. Secondary objectives included pharmacokinetics, changes in selected electrocardiogram (ECG) parameters, and categorical changes to the QTcF, PR, and QRS intervals and heart rate (pooled analysis). Thirty-two participants were randomized in A2107 and 56 in X2101; demographic data were similar across treatment groups. The estimated placebo-adjusted change from Day 1 baseline in QTcF at the geometric mean maximum drug concentration was 1.61 (90% confidence intervals: 0.39 to 2.82), 1.26 (-0.42 to 2.94), and 0.84 (-1.54 to 3.22) ms for 400, 800, and 1200 mg doses, respectively. These primary results are consistent with no risk of QT prolongation at single, supratherapeutic, oral iptacopan doses; secondary ECG data support this conclusion. No deaths, serious adverse events (AEs), or AEs leading to study discontinuation were reported; iptacopan was overall well tolerated. A high systemic exposure of iptacopan was confirmed, allowing us to conclude that single, supratherapeutic, oral doses of iptacopan had no QTcF prolongation or proarrhythmic potential in healthy individuals.
To assess cardiac and safety parameters of iptacopan (an oral, selective, reversible, small-molecule factor B inhibitor), we conducted a phase I, single ascending dose (SAD), exposure-response study (A2107) instead of a traditional thorough QT study. Healthy participants were randomized 3:1 to receive a single, supratherapeutic, oral dose of iptacopan 400, 800, or 1200 mg, or placebo. A2107's primary objectives were to assess iptacopan's effect on Fridericia-corrected QT interval (QTcF) in a pooled analysis with the SAD phase of the first-in-human X2101 study, and the safety and tolerability of supratherapeutic iptacopan doses in participants. Secondary objectives included pharmacokinetics, changes in selected electrocardiogram (ECG) parameters, and categorical changes to the QTcF, PR, and QRS intervals and heart rate (pooled analysis). Thirty-two participants were randomized in A2107 and 56 in X2101; demographic data were similar across treatment groups. The estimated placebo-adjusted change from Day 1 baseline in QTcF at the geometric mean maximum drug concentration was 1.61 (90% confidence intervals: 0.39 to 2.82), 1.26 (-0.42 to 2.94), and 0.84 (-1.54 to 3.22) ms for 400, 800, and 1200 mg doses, respectively. These primary results are consistent with no risk of QT prolongation at single, supratherapeutic, oral iptacopan doses; secondary ECG data support this conclusion. No deaths, serious adverse events (AEs), or AEs leading to study discontinuation were reported; iptacopan was overall well tolerated. A high systemic exposure of iptacopan was confirmed, allowing us to conclude that single, supratherapeutic, oral doses of iptacopan had no QTcF prolongation or proarrhythmic potential in healthy individuals.
More Related Videos
 07:35
Assessment of Open Probability of the Mitochondrial Permeability Transition Pore in the Setting of Coenzyme Q Excess
Published on: June 1, 2022
 08:28
Methods for ECG Evaluation of Indicators of Cardiac Risk, and Susceptibility to Aconitine-induced Arrhythmias in Rats Following Status Epilepticus
Published on: April 5, 2011
Related Concept Videos
 01:15
Determination of Multiple Dosing Parameters: Steady-State, Minimum and Maximum Concentrations
 01:20
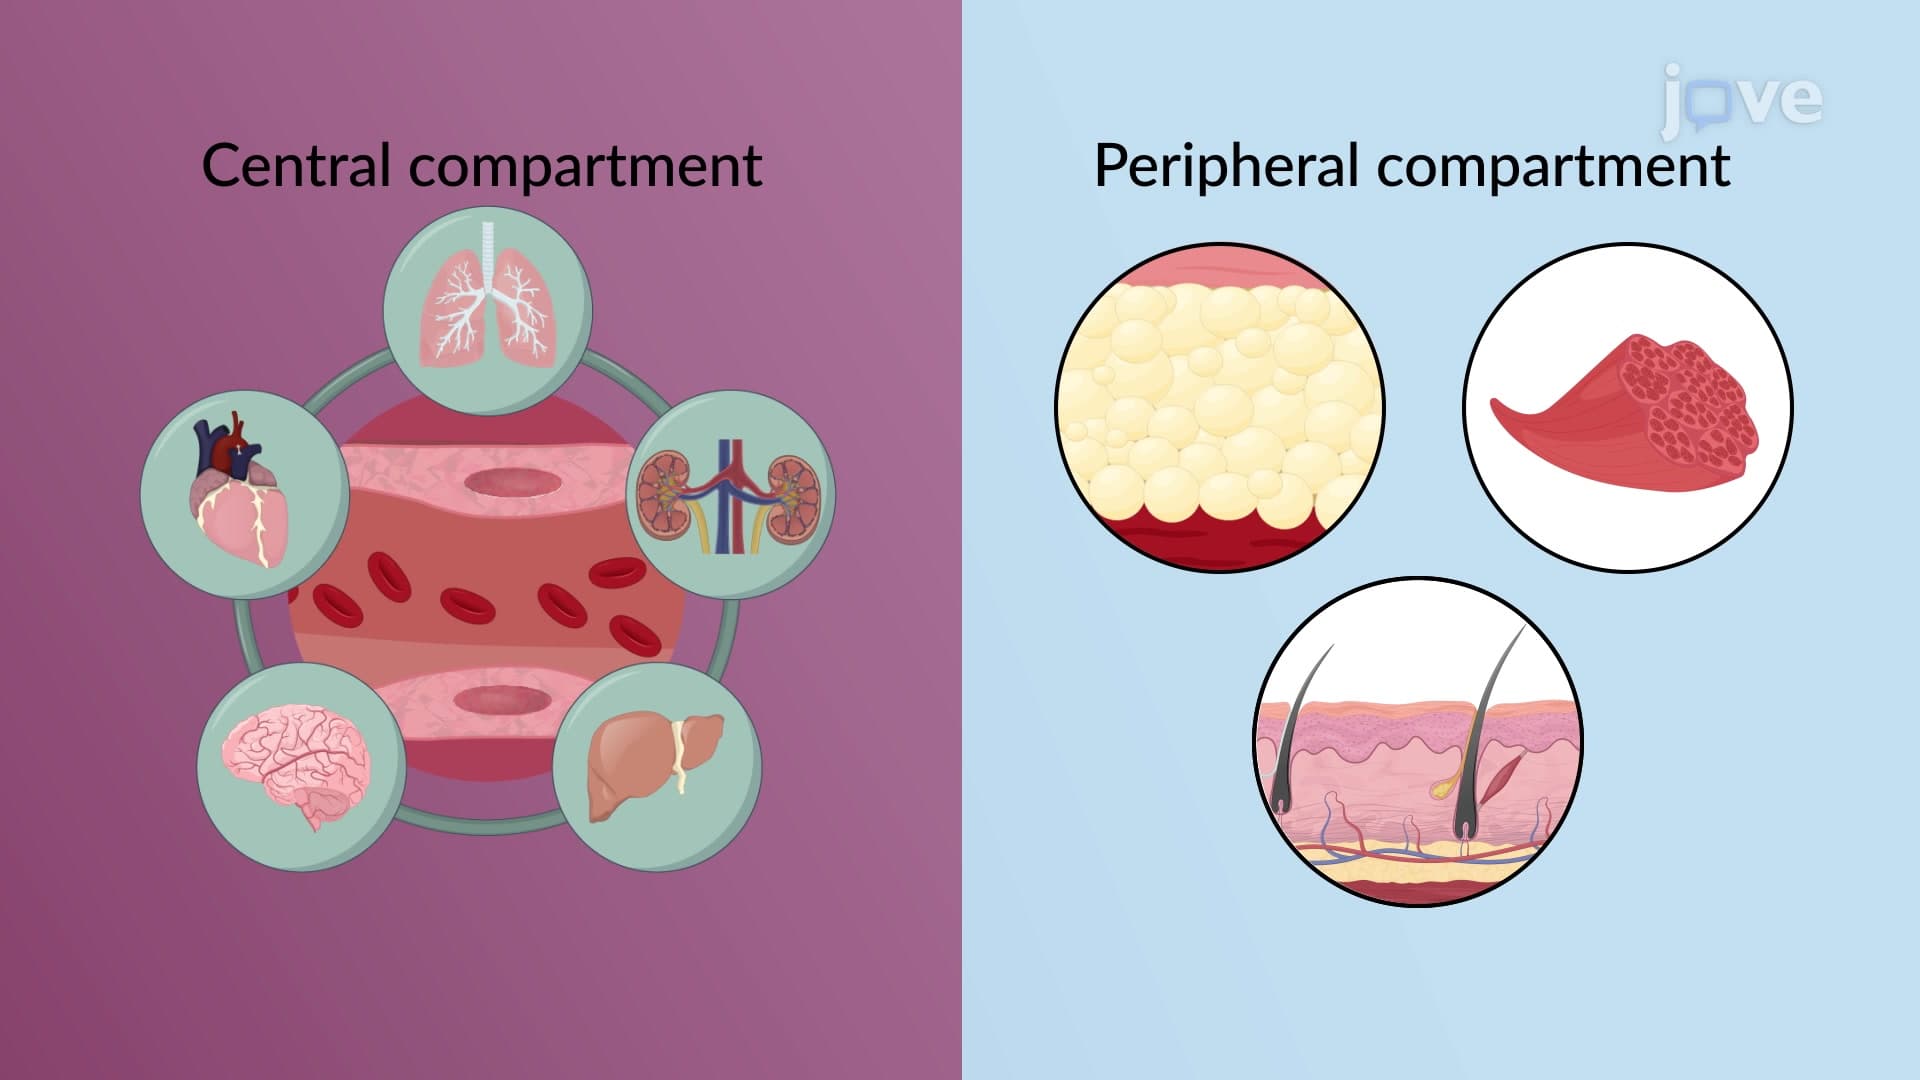
Compartment Models: Two-Compartment Model
 01:15
Drug Concentration Versus Time Correlation
Two pivotal parameters are the minimum effective concentration (MEC) and the minimum toxic concentration (MTC). The MEC is the...
 01:19
One-Compartment Open Model for IV Bolus Administration: General Considerations
The drug's presence in the body is defined by an equation representing the difference between the rates of drug entry and exit. Key parameters—elimination rate constant,...
 00:54
Noncompartmental Analysis: Miscellaneous Pharmacokinetic Parameters
One key aspect of the noncompartmental approach is determining a drug's total clearance. This can be done by dividing the drug dose by the area under the concentration-time curve from zero to infinity. The area under the concentration-time curve represents the drug's...
 00:59
Model-Independent Approaches for Pharmacokinetic Data: Noncompartmental Analysis
One important characteristic of noncompartmental analyses is that drug exposure increases proportionally with increasing doses. This...