Related Experiment Video
Updated: Mar 28, 2026


Analyzing Melts and Fluids from Ab Initio Molecular Dynamics Simulations with the UMD Package
Published on: September 17, 2021
Molecular dynamics simulation of nitrogen diffusion in iron and iron nitrides using ab initio data trained machine
Peijie Feng1, Aditya Dilip Lele2,3, Minhyeok Lee1
1Department of Mechanical Engineering, The University of Tokyo, 7-3-1 Hongo, Bunkyo-ku, Tokyo 113-8656, Japan. ysuzuki@mesl.t.u-tokyo.ac.jp.

Abstract:
Ammonia is receiving increasing attention as a hydrogen energy carrier and a green fuel for achieving carbon neutrality. Combustors utilizing ammonia as a fuel are subject to "unwanted" surface nitriding, which hardens and embrittles the metal walls of combustors. Predicting the nitridation rate and nitride layer thicknesses with Fick's diffusion law requires temperature- and concentration-dependent nitrogen atoms' diffusion coefficients in the relevant iron and iron nitrides. Experimentally determining the diffusion coefficients is challenging due to steep N-gradient, and concurrent phase transformations. In this study, we calculated the nitrogen atom diffusion coefficients using molecular dynamics (MD) driven by a machine learning interatomic potential (MLP). The MLP was trained on snapshots extracted from ab initio molecular dynamics (AIMD) trajectories covering different temperatures and nitrogen atom concentrations, and the contribution of each subset to the model's accuracy was quantified. The results show that the MLP reproduces independently the calculated diffusion energy barriers obtained using density functional theory (DFT) in α-, γ-iron and γ'-, ε-iron nitrides. For ε-iron nitride, the MLP-driven MD-computed self-diffusion coefficient was converted into chemical (Fickian) diffusion coefficients, enabling direct comparison with experiments. Our MLP-driven MD simulation delivers diffusion coefficients approaching ab initio accuracy, enabling precise concentration- and temperature-resolved modeling of nitrogen diffusion in iron and its nitrides. Arrhenius fitting to the 830-1500 K chemical diffusion coefficients reproduce the experimental activation energy and pre-exponential factor. Extrapolation to 823 K, the temperature at which most of the experiments are performed, yields a coefficient that falls within the experimental uncertainty. The present MLP-MD model will facilitate the mechanistic understanding of iron surface nitridation and quantitative predictions of "unwanted" nitriding of iron-based combustor wall materials induced by ammonia at high temperatures.
More Related Videos
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
 11:29
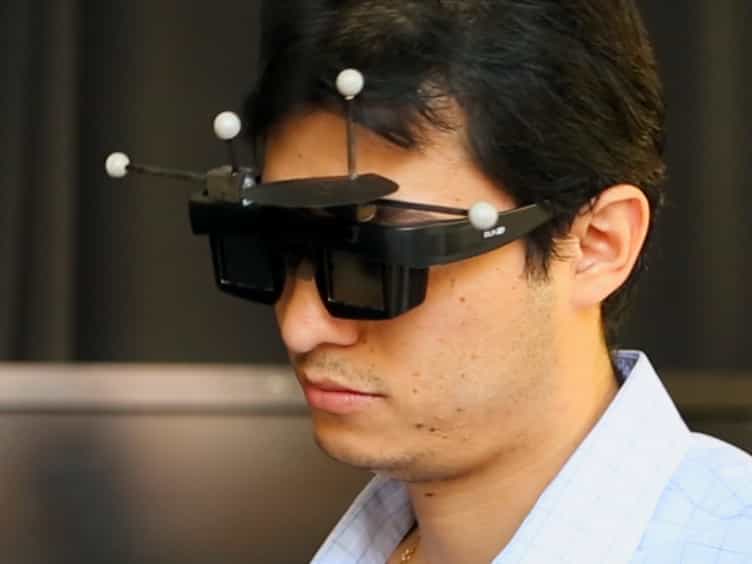
Novel 3D/VR Interactive Environment for MD Simulations, Visualization and Analysis
Published on: December 18, 2014
Related Concept Videos
 01:22
Inorganic Nitrogen Assimilation
 03:48
Behavior of Gas Molecules: Molecular Diffusion, Mean Free Path, and Effusion
 01:49
The Nitrogen Cycle
 01:31
Ferromagnetism