Related Experiment Video
Updated: Sep 19, 2026

Modeling Ligands into Maps Derived from Electron Cryomicroscopy
Published on: July 19, 2024
Structure-based molecular modeling of a novel S-Remiketamine scaffold and derived arylcyclohexanone analogues: NMDA
Ahmed A A M M Alkhatip1,2, Usama Raza3, Ehab Farag2
1Department of Anaesthesia, Birmingham Children's Hospital, Birmingham, United Kingdom.

Introduction:
Short-acting ketamine-related scaffold design remains an important medicinal-chemistry problem, but computational studies require conservative interpretation until receptor, enzymatic, and pharmacokinetic validation is available.
Methods:
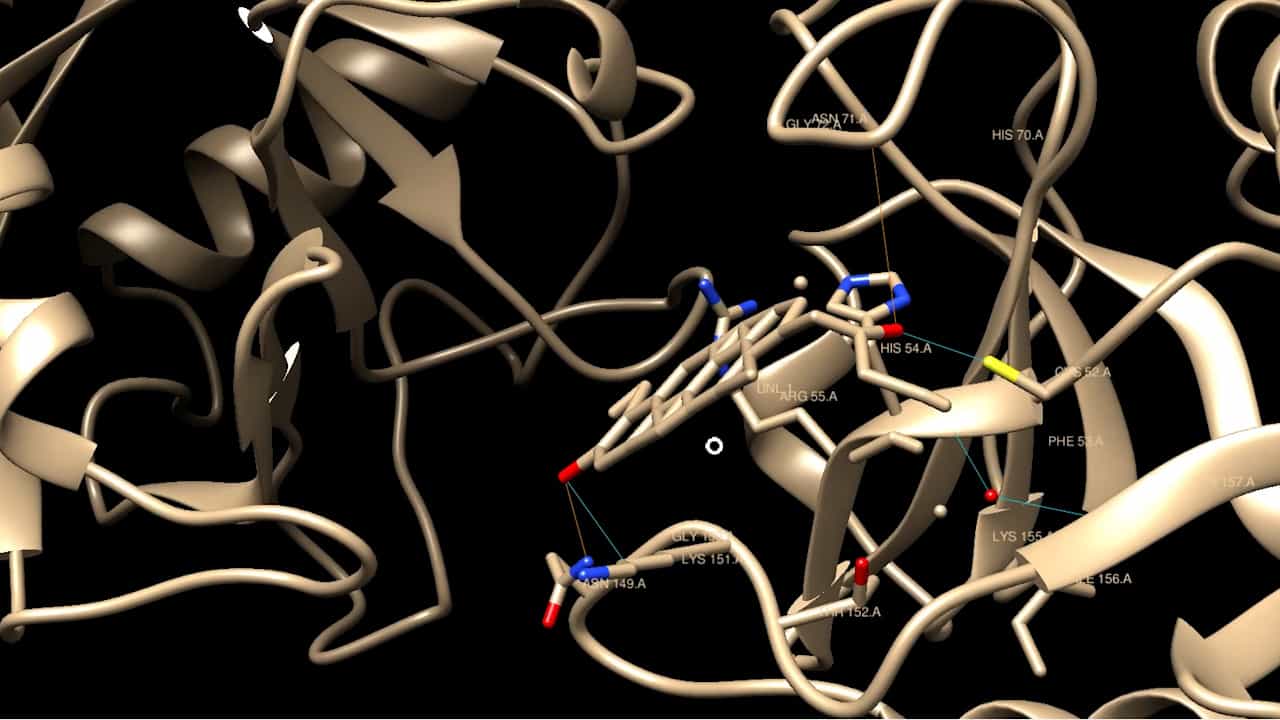
S-ketamine, S-Remiketamine, and R1-R20 analogues were assessed using ligand curation, NMDA receptor docking, cross-target docking, explicit-solvent molecular dynamics, MM-GBSA analysis, CES1-oriented geometric assessment, and predictive ADMET/metabolite-route analysis.
Results:
Static NMDA docking ranked R2 highest (Glide score -7.308 kcal/mol), while S-Remiketamine showed a more favorable predicted NMDA score than S-ketamine (-6.448 vs. -5.981 kcal/mol). Within the dynamically evaluated subset, NMDA end-state MM-GBSA values were -41.53 kcal/mol for R2, -28.80 kcal/mol for R11, and -42.10 kcal/mol for R14. In CES1 modeling, R14 was the only evaluated compound whose MM-GBSA became more favorable from 0 to 100 ns (-83.91 to -99.44 kcal/mol).
Discussion:
These results support R2 as the strongest static NMDA docking/NMDA-preference benchmark and R14 as the most internally consistent integrated computational candidate. The findings are hypothesis-generating and require synthesis and experimental validation.