Related Experiment Video
Updated: Oct 25, 2026
Isolating Free Carbenes, their Mixed Dimers and Organic Radicals
Published on: April 19, 2019
When carbonyls beat aromaticity: an additive framework for predicting tautomeric preferences
Lucas Araujo1, Felipe Fantuzzi2, Pierre M Esteves1
1Instituto de Química, Universidade Federal do Rio de Janeiro Av. Athos da Silveira Ramos, 149, CT, A-622, Cid. Univ. Rio de Janeiro RJ 21941-909 Brazil thiago@iq.ufrj.br pesteves@iq.ufrj.br.

Abstract:
Aromatic structures are often regarded as privileged stabilising motifs, generally retained even when competing structural effects are present. Here, we show that functional-group energetics can overturn this expectation in heteroaromatic tautomerism. Analysis of the 2-hydroxypyridine/2-pyridone equilibrium reveals a near-cancellation between carbonyl stabilisation and the energetic penalty associated with perturbing the aromatic ring framework, accounting for the small experimental energy gap. Extending this approach to pyridines, diazines, and triazines reveals a simple additive pattern: hydroxy/carbonyl exchange favours carbonylated, non-aromatic tautomers, typically requiring two C[double bond, length as m-dash]O/N-H units to invert the preference, whereas amino/imine and methyl/methylidene exchanges reinforce aromatic tautomers. Spin-coupled generalised valence bond (SCGVB) calculations analysed within the GPF-EP method show that the quasi-classical component favours the aromatic 6c-6e π framework, whereas quantum interference favours localised 2c-2e bonds, especially C[double bond, length as m-dash]O. This provides an energy-decomposition expression of π-distortivity: without interference, the aromatic tautomers would predominate across all series. Complementary NBO analysis suggests that this trend reflects stronger lactam-type donor-acceptor delocalisation in carbonylated N-heteroaromatic tautomers than in the corresponding imine, methylidene, or all-carbon benzenoid keto analogues. The model rationalises benzenoid phenols as boundary cases and predicts tautomeric preferences along the deamination pathway from melamine to cyanuric acid. These results establish a framework for predicting tautomeric preferences and clarify how aromatic-ring stabilisation is balanced against functional-group energetics.
Related Concept Videos
 01:20
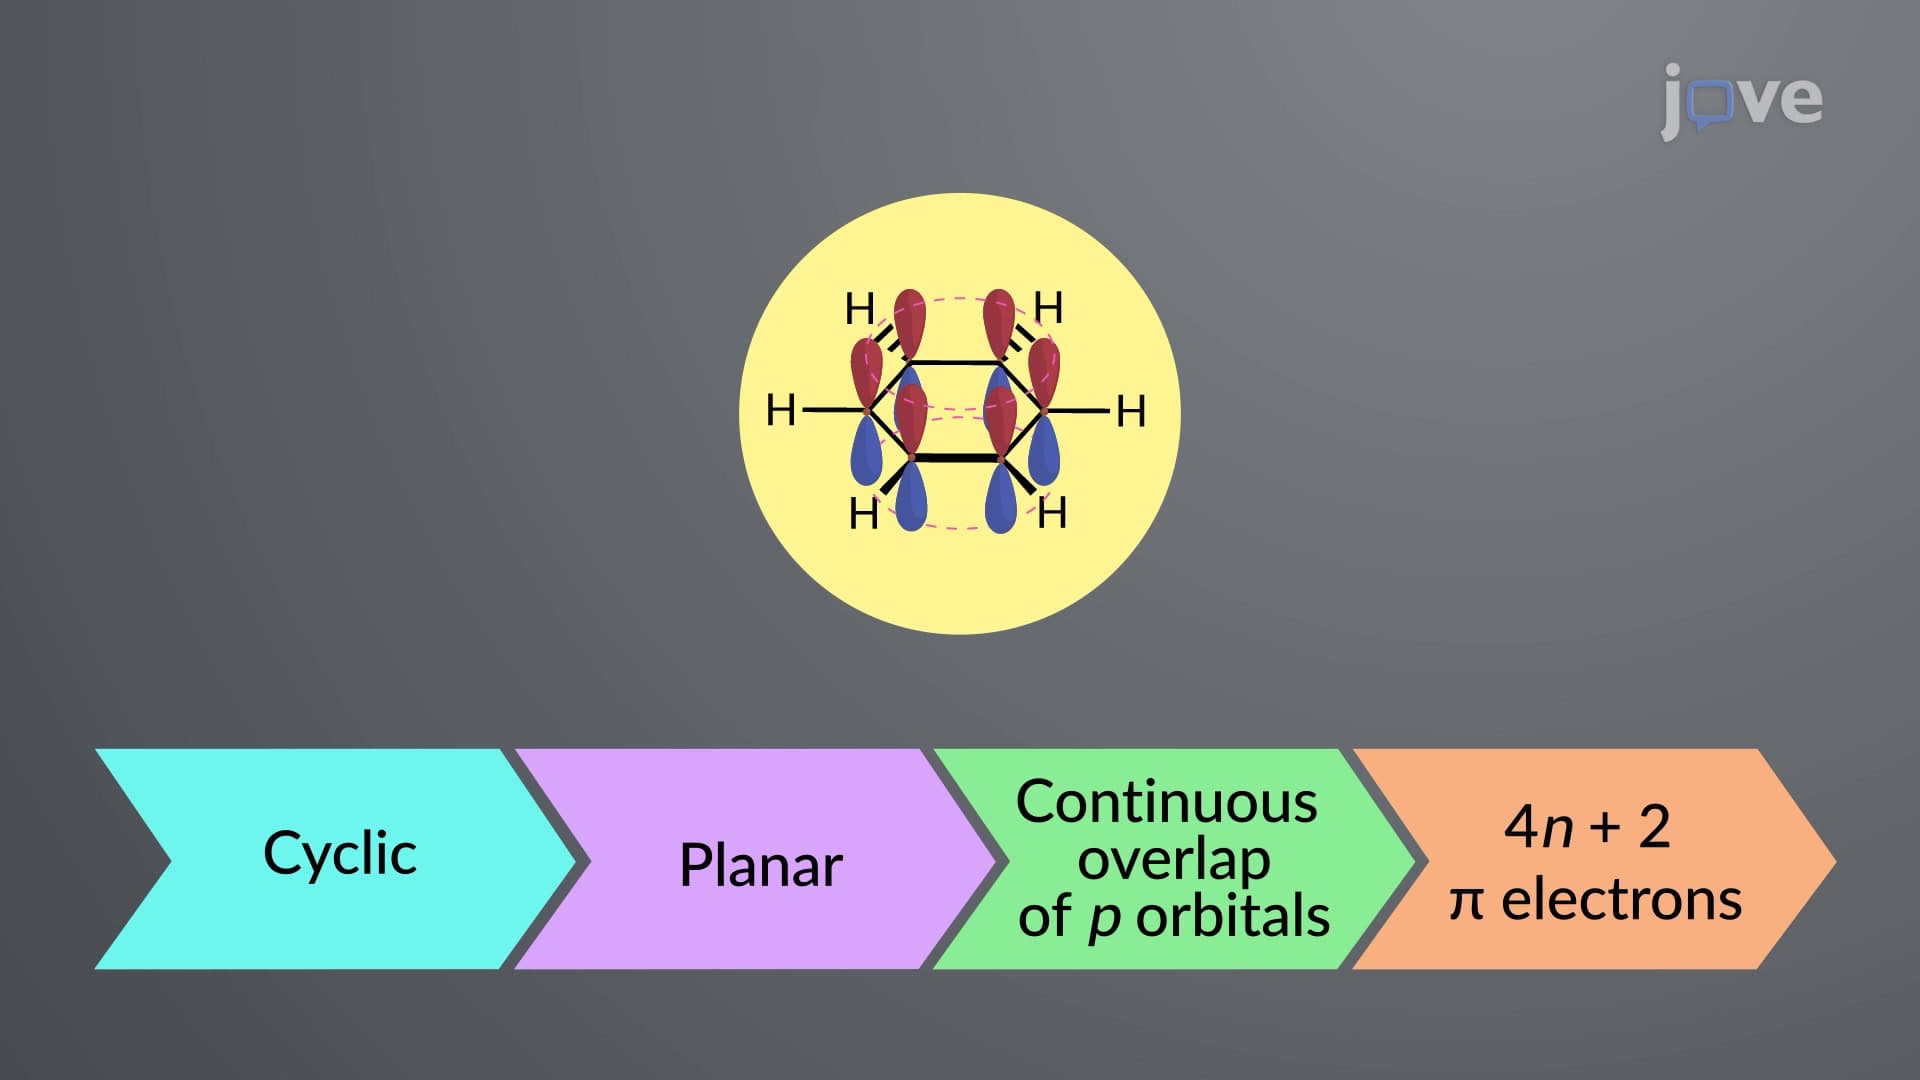
Criteria for Aromaticity and the Hückel 4n + 2 Rule
For the first time, Eric Hückel, a German chemical physicist, derived a set of structural features for a compound to be classified as aromatic. This is now known as Hückel’s rule or the 4n + 2 rule.
 01:18
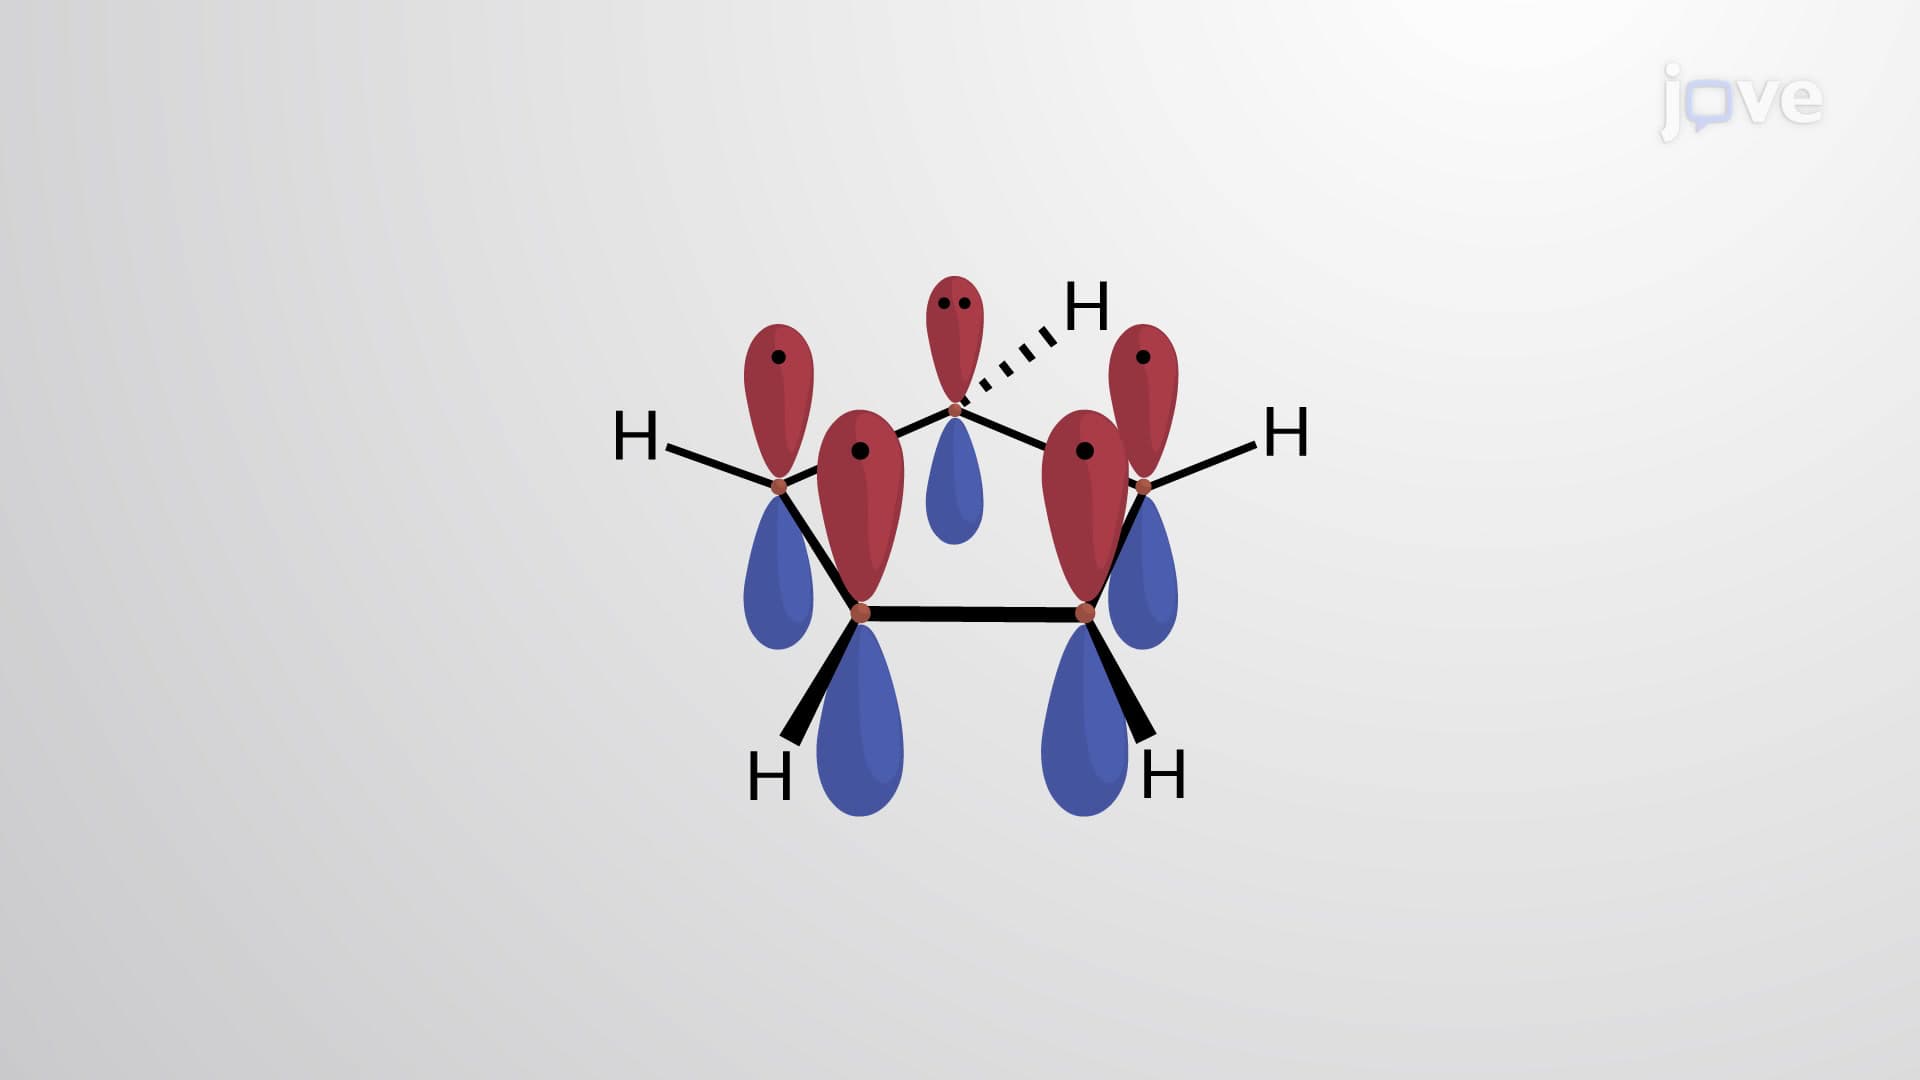
Aromatic Hydrocarbon Anions: Structural Overview
Due to the absence of continuous overlap of p...
 01:25
Aromatic Compounds: Overview
In 1825, Faraday isolated benzene...
 01:18
Frost Circles for Different Conjugated Systems
 01:18
Aromatic Hydrocarbon Cations: Structural Overview
Removing one hydrogen from the intervening CH2 group with both...
 01:13
Relative Reactivity of Carboxylic Acid Derivatives
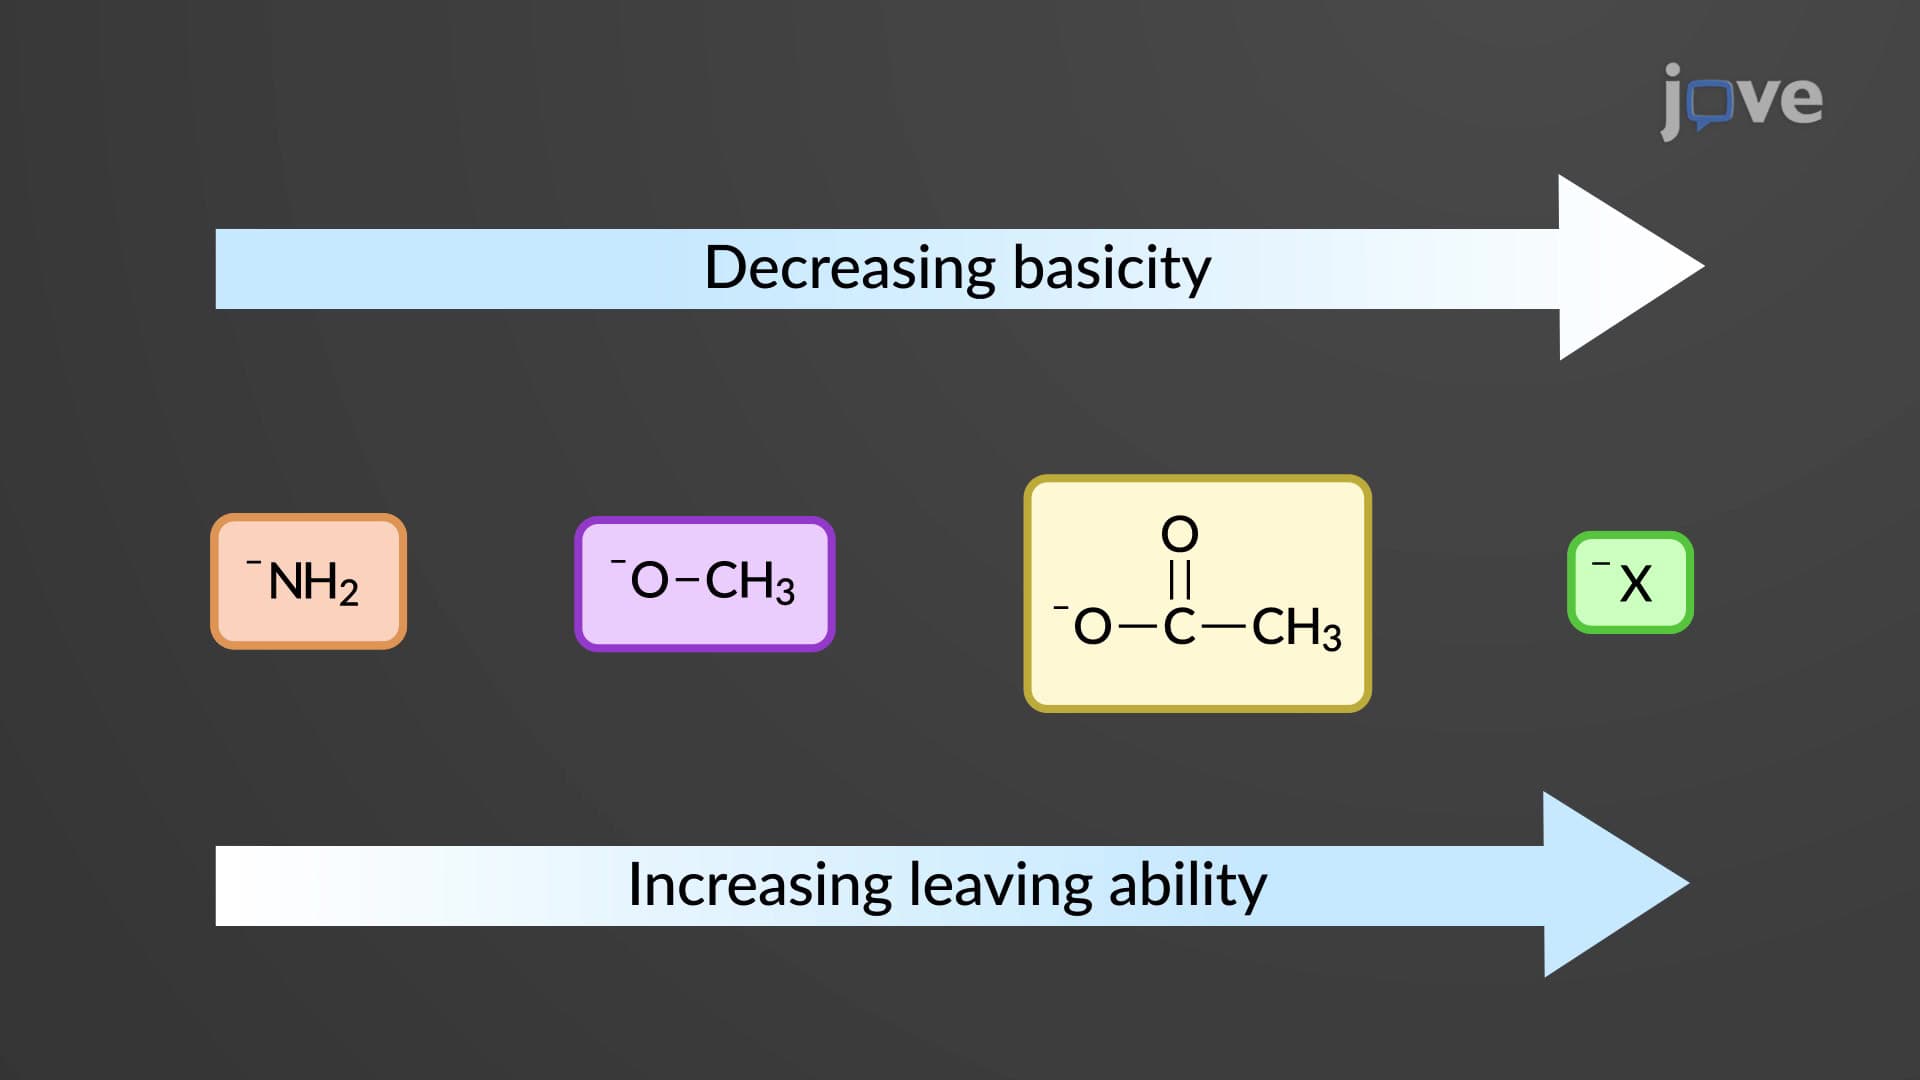
A key factor in assessing the reactivity of the acid derivatives is the basicity of the substituent or the leaving group. The lower the basicity of the leaving group, the higher the reactivity of the derivative. The basicity of the leaving group follows this order:
Halide ions < Acyloxy ions < Alkoxy ions < Amine ions