Related Experiment Video
Updated: Oct 26, 2026
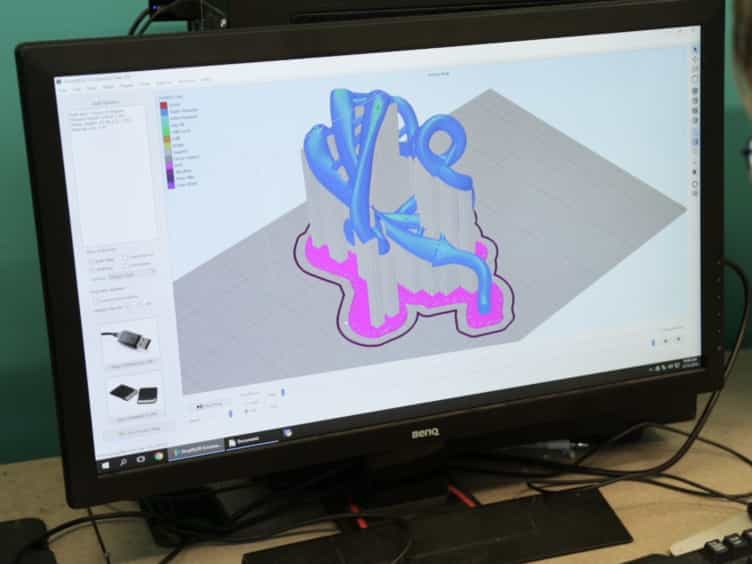
3D Printing of Biomolecular Models for Research and Pedagogy
Published on: March 13, 2017

Abstract:
AlphaFold 3 predicts protein structures with remarkable accuracy, yet how structural information emerges within the model remains poorly understood. Here, through causal interventions on internal representations and direct probing of every Pairformer block, we trace the formation of global protein geometry and identify the multiple sequence alignment (MSA) as a structural shortcut to the fold. Removing the MSA largely preserves local secondary structure while disrupting the long-range relationships that define global topology. Restoring the MSA-enriched pair representation at only forty residues recovers most of this lost organization, including at pairs never directly modified. This contribution depends on the detailed direction of the MSA module's output rather than its magnitude. The Pairformer rapidly converts this signal into global geometry: the final fold becomes recoverable by approximately block 9 of 48 for a majority of proteins, roughly twenty-seven blocks before the model's decoder can render it, whereas without the MSA it remains inaccessible for most proteins throughout the pass. Which homologs are supplied shapes this trajectory more strongly than which query is supplied; it persists for a designed query that never evolved but collapses for a shuffled sequence. Most importantly, an alignment built for a different protein that shares the fold, supplied only at the structurally corresponding columns, raises the median TM-score against experiment from 0.44 to 0.72, while the same alignment shifted a few residues along the chain performs worse than supplying no alignment at all. What AlphaFold 3 reads from an alignment is therefore a description of the fold itself, transferable between proteins that share one, rather than the query's own evolutionary history. This explains both its accuracy and the limits of what it has solved.
More Related Videos
 10:01
Structure-Guided Design and Development of Novel Cyclophilin A Inhibitors and Ganoderiol-F Derivatives: An In-Silico Approach
Published on: June 23, 2026
 10:29
Quantitative Structure-Activity Relationship, Activity Prediction, and Molecular Dynamics of Non-nucleotide Reverse Transcriptase Inhibitors
Published on: May 9, 2025
Related Concept Videos
 01:04
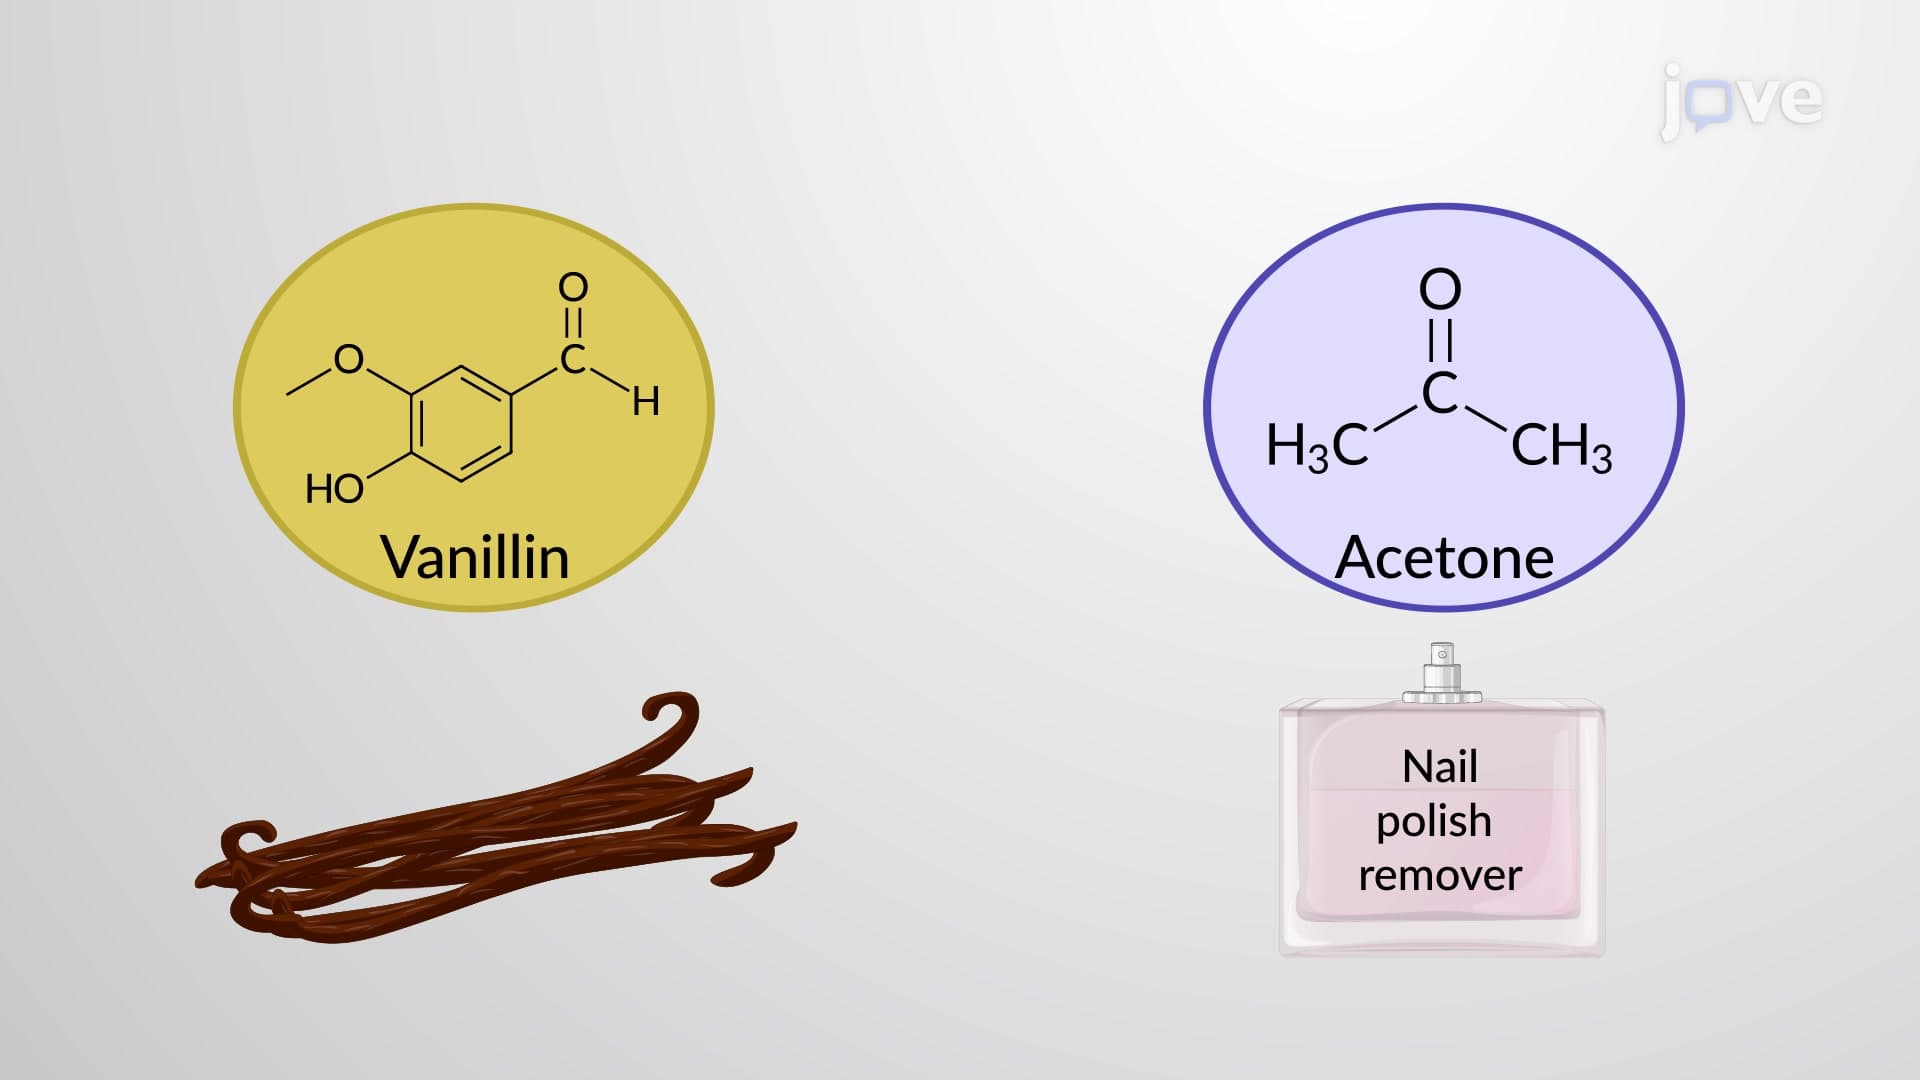
Structures of Aldehydes and Ketones
In aldehydes (Figures 1a and 1b), the carbonyl...
 02:18
Fischer Projections
 01:24
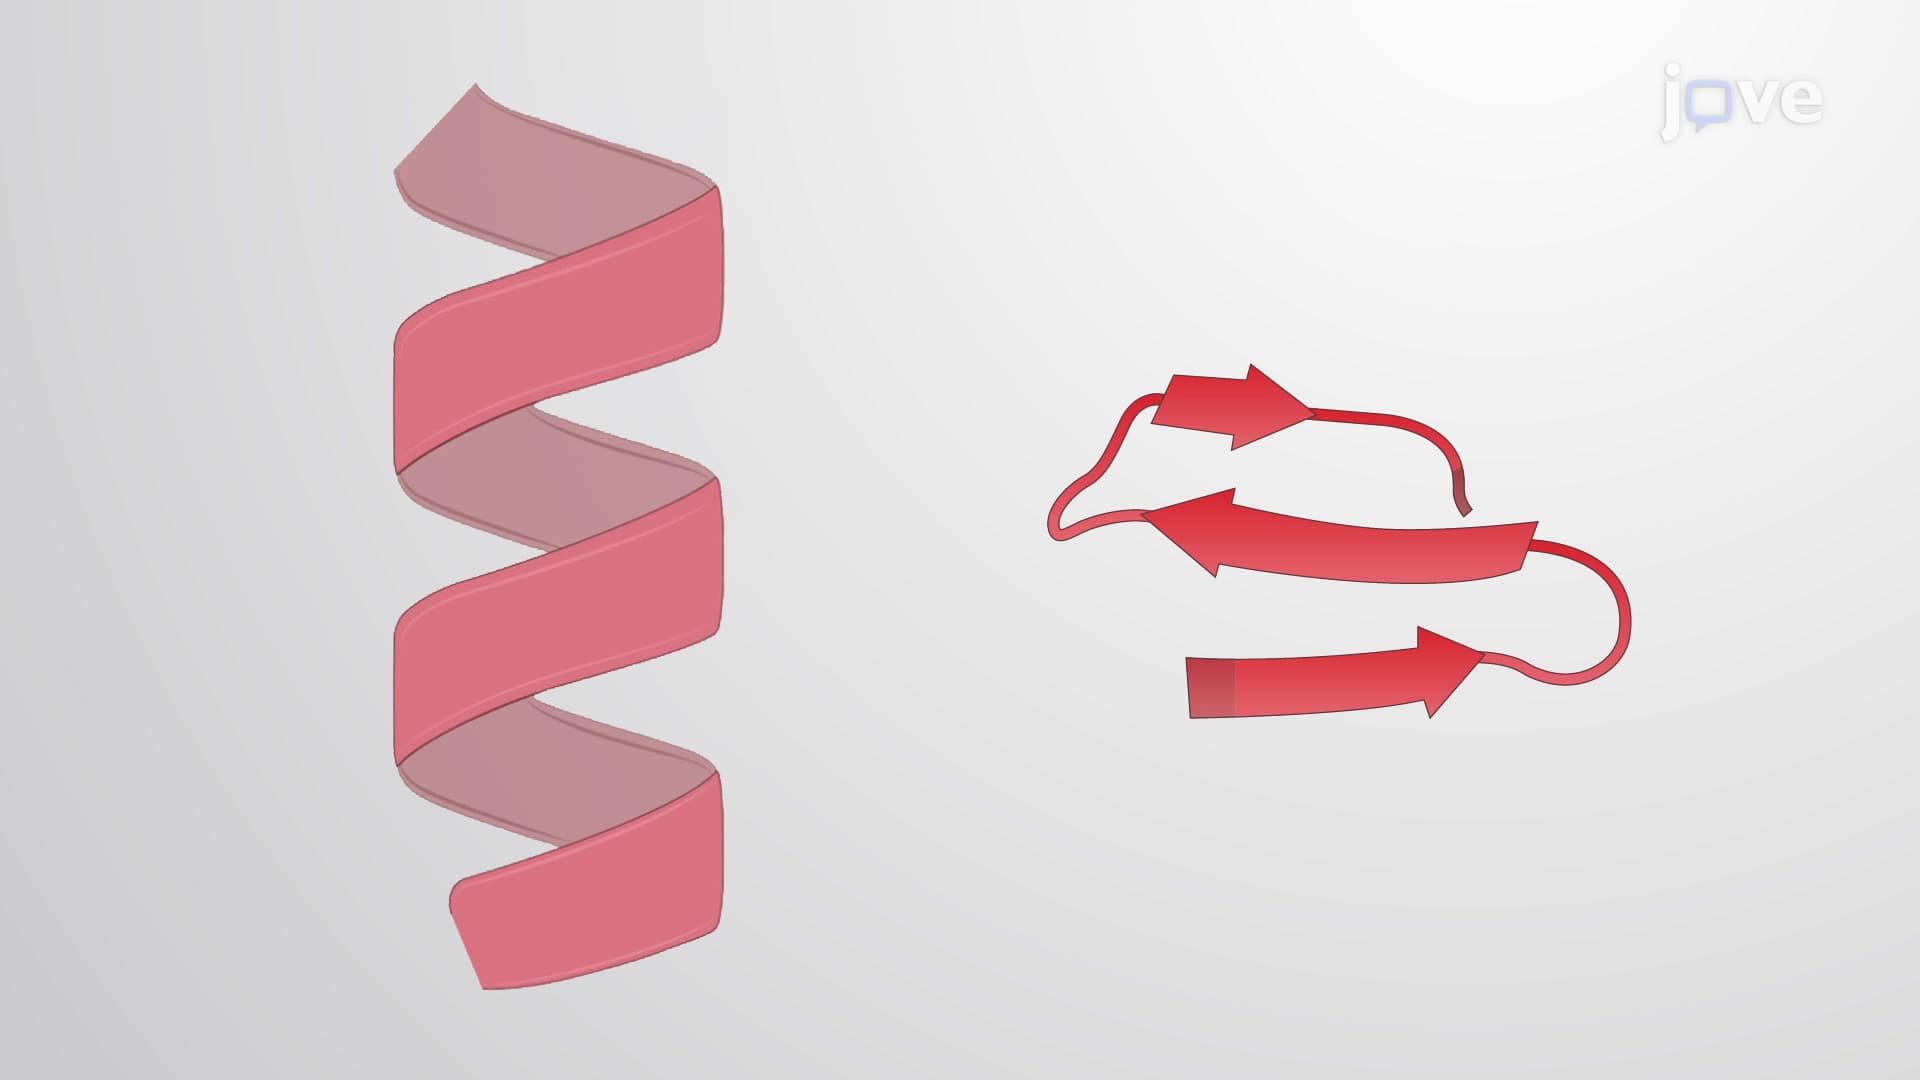
Protein Organization
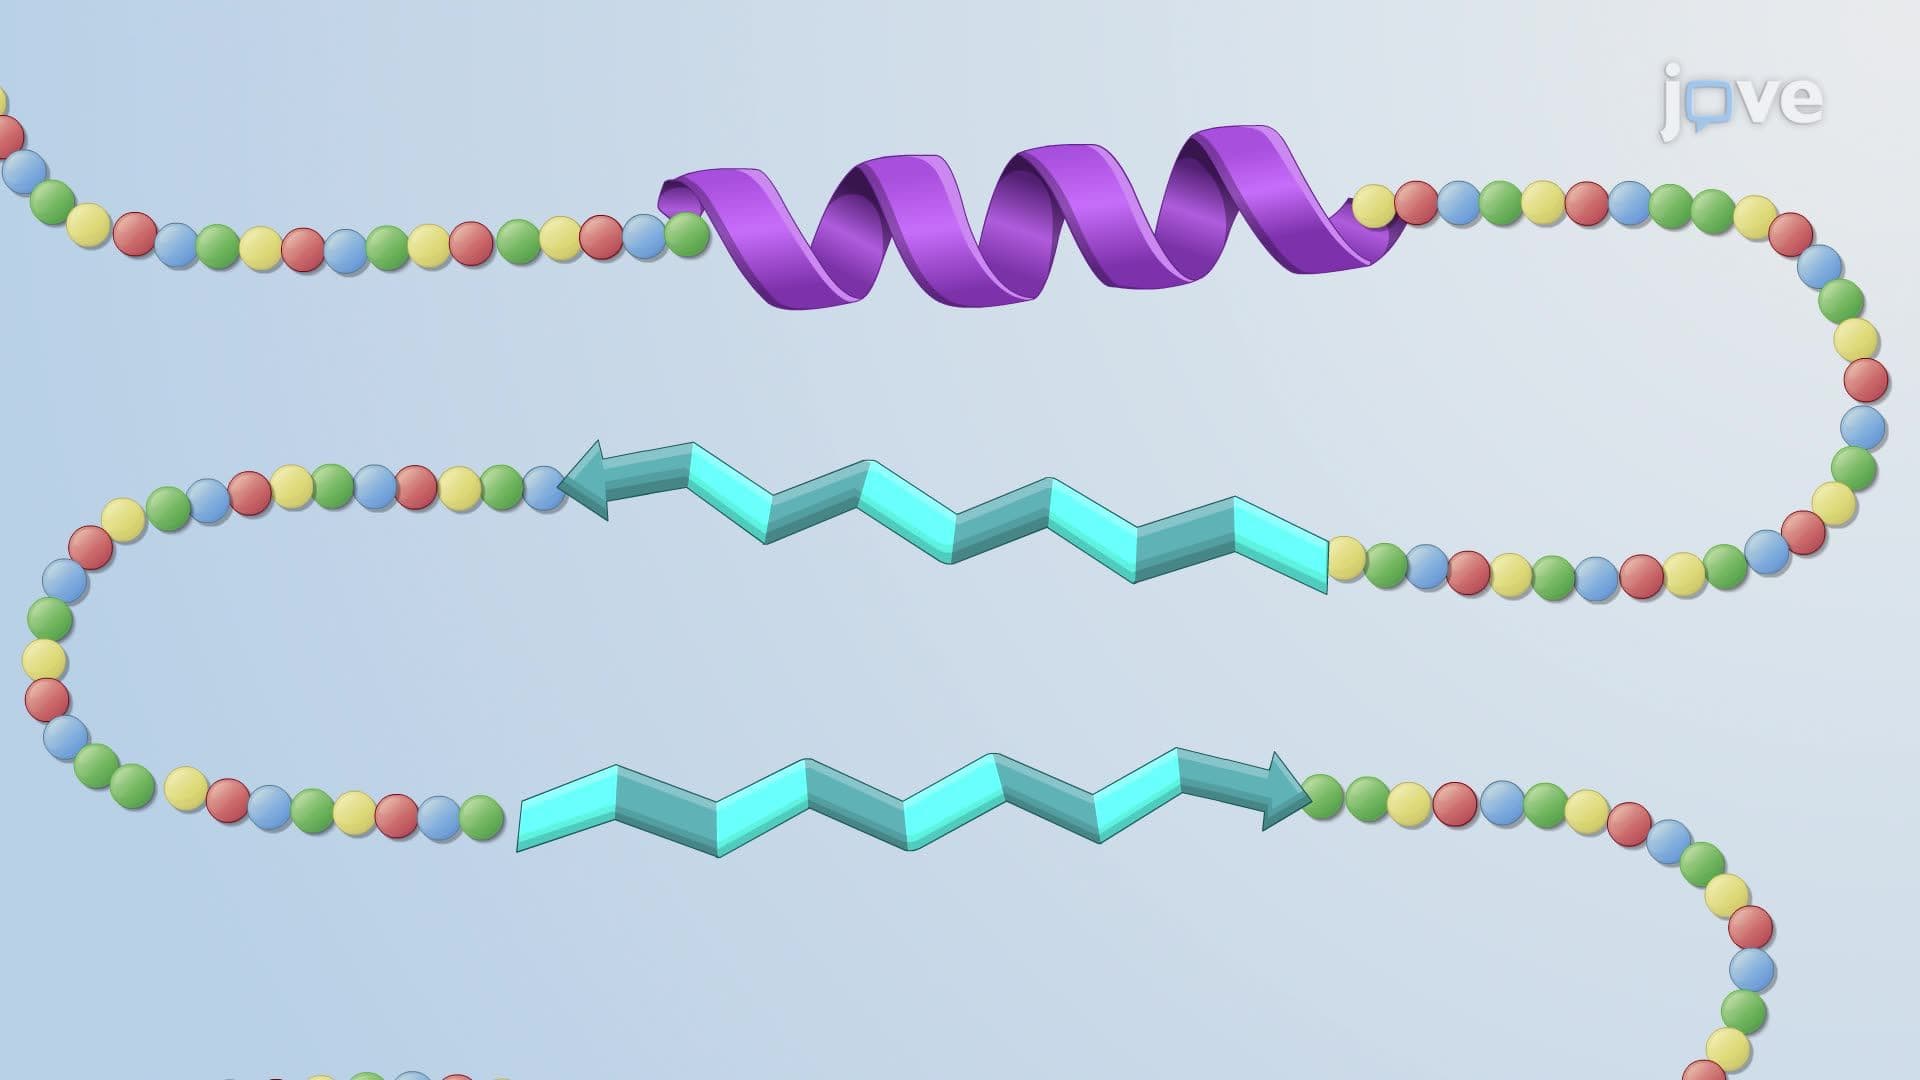
The primary structure of a protein is its amino acid sequence.
 01:28
Protein Organization
01:24Protein Organization
The primary structure of a protein is its amino acid sequence.
 02:47
Structure and Bonding of Alkenes
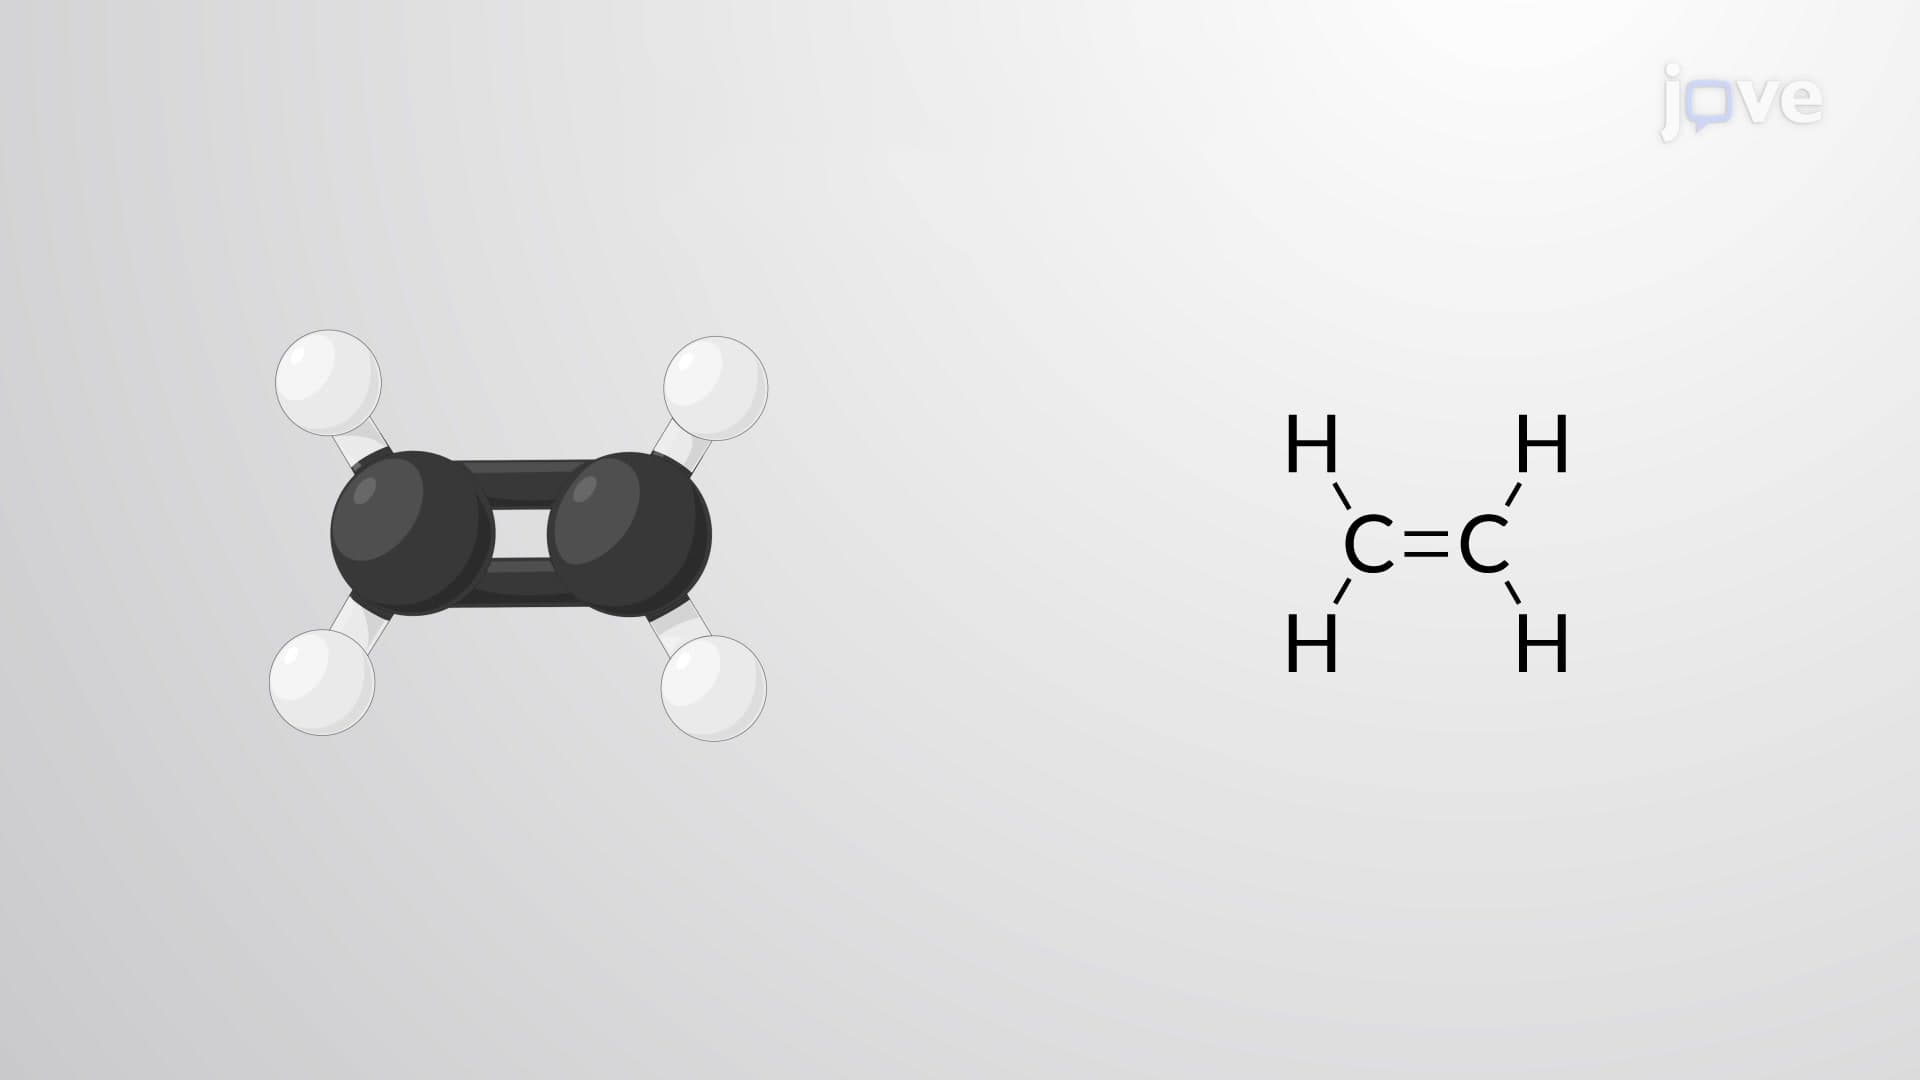
Doubly bonded carbons are sp2 hybridized and have a trigonal planar geometry. The double bond is composed of a σ bond formed by the overlap of hybrid orbitals and a π bond produced by the lateral overlap of unhybridized 2p orbitals on both the carbons. Each carbon atom is bonded to two hydrogen atoms...