Related Experiment Video
Updated: Aug 13, 2026

Mass Spectrometry and Luminogenic-based Approaches to Characterize Phase I Metabolic Competency of In Vitro Cell Cultures
Published on: March 28, 2017
Structure-activity relationships for triphenylethylene antiestrogens on hepatic phase-I and phase-II enzyme
E F Nuwaysir1, Y P Dragan, R McCague
1Environmental Toxicology Center, University of Wisconsin, Madison, USA.

Abstract:
To better understand the mechanism(s) by which tamoxifen induces rat hepatic CYPIIB2 and suppresses GSTA1, structure-activity studies were performed. Compounds employed in these studies included: tamoxifen, fixed-ring tamoxifen, ethylated fixed-ring tamoxifen, pyrrolidino-tamoxifen, 4-iodotamoxifen, idoxifene, and toremifene. With respect to GSTA1 suppression, tamoxifen, fixed-ring tamoxifen, 4-iodotamoxifen, idoxifene, and toremifene were all potent suppressors of GSTA1, while ethylated fixed-ring tamoxifen and pyrrolidino-tamoxifen were completely without activity. The results suggest that the aminoethoxy side chain plays a crucial role in GSTA1 suppression, and that 4-iodination may potentiate this activity. With respect to induction of CYPIIB2, tamoxifen, fixed-ring tamoxifen, and ethylated fixed-ring tamoxifen were inducers of this enzyme, while toremifene and 4-iodotamoxifen were inactive, suggesting that the aminoethoxy side chain is not a structural determinant of CYPIIB2 induction. Because ethylated fixed-ring tamoxifen, toremifene, and 4-iodotamoxifen had differential activities in the two assays, we conclude that CYPIIB2 induction and GSTA1 suppression by triphenylethylenes are the result of two separate and distinct mechanistic pathways. Structure-activity relationships for GSTA1 suppression and CYPIIB2 induction were compared with previously published relationships for triphenylethylene: 1) estrogen receptor relative binding affinity; 2) calmodulin antagonism; 3) antiuterotrophic activity; and 4) antagonism of MCF-7 cell growth. No clear correlation was observed between the effects on CYPIIB2 and these other four activities, suggesting no relationship between the mechanisms responsible for these effects. Similarly, no precise correlation was observed between GSTA1 suppression and these other activities, although rough similarities were observed for relative binding affinity and antiuterotrophic activity. This suggests that the mechanisms responsible for CYPIIB2 induction and GSTA1 suppression are not related to the mechanisms of action for these other documented activities, and may represent different mechanistic pathways.
Insights
Tamoxifen
Area of Science:
- Pharmacology and Toxicology
- Enzyme Kinetics
- Drug Metabolism
Background:
- Tamoxifen is a widely used drug with known effects on liver enzymes.
- Understanding the precise mechanisms of tamoxifen's action is crucial for optimizing its therapeutic use and predicting potential drug interactions.
Purpose of the Study:
- To elucidate the distinct mechanisms by which tamoxifen and its analogs induce rat hepatic CYPIIB2 and suppress glutathione S-transferase alpha 1 (GSTA1).
- To investigate the structure-activity relationships governing these two distinct biological effects.
Main Methods:
- Synthesis and testing of a series of tamoxifen analogs, including fixed-ring tamoxifen, ethylated fixed-ring tamoxifen, pyrrolidino-tamoxifen, 4-iodotamoxifen, idoxifene, and toremifene.
- Assessing the ability of these compounds to induce CYPIIB2 and suppress GSTA1 in rat liver models.
- Comparing the observed structure-activity relationships with known activities such as estrogen receptor binding and calmodulin antagonism.
Main Results:
- The aminoethoxy side chain is critical for GSTA1 suppression, with 4-iodination potentially enhancing this activity.
- CYPIIB2 induction is not dependent on the aminoethoxy side chain, as demonstrated by the differential activity of various analogs.
- Differential activities of analogs in CYPIIB2 induction and GSTA1 suppression confirm distinct mechanistic pathways for these effects.
Conclusions:
- Tamoxifen-induced CYPIIB2 induction and GSTA1 suppression in rats operate via separate and distinct mechanistic pathways.
- The mechanisms underlying CYPIIB2 induction and GSTA1 suppression by triphenylethylenes are unrelated to estrogen receptor binding, calmodulin antagonism, antiuterotrophic activity, or MCF-7 cell growth antagonism.
More Related Videos
 09:33
Formation of Covalent DNA Adducts by Enzymatically Activated Carcinogens and Drugs In Vitro and Their Determination by 32P-postlabeling
Published on: March 20, 2018
 08:59
An Intestine/Liver Microphysiological System for Drug Pharmacokinetic and Toxicological Assessment
Published on: December 3, 2020
Related Concept Videos
 01:17
Drug Metabolism: Phase I Reactions
 01:14
Drug Metabolism: Phase II Reactions
 01:19
Phase I Reactions: Oxidation of Aliphatic and Aromatic Carbon-Containing Systems
Oxidation reactions are fundamental in aromatic carbon-containing systems. An example is the hydroxylation of phenobarbital, a process that transforms it into...
 01:19
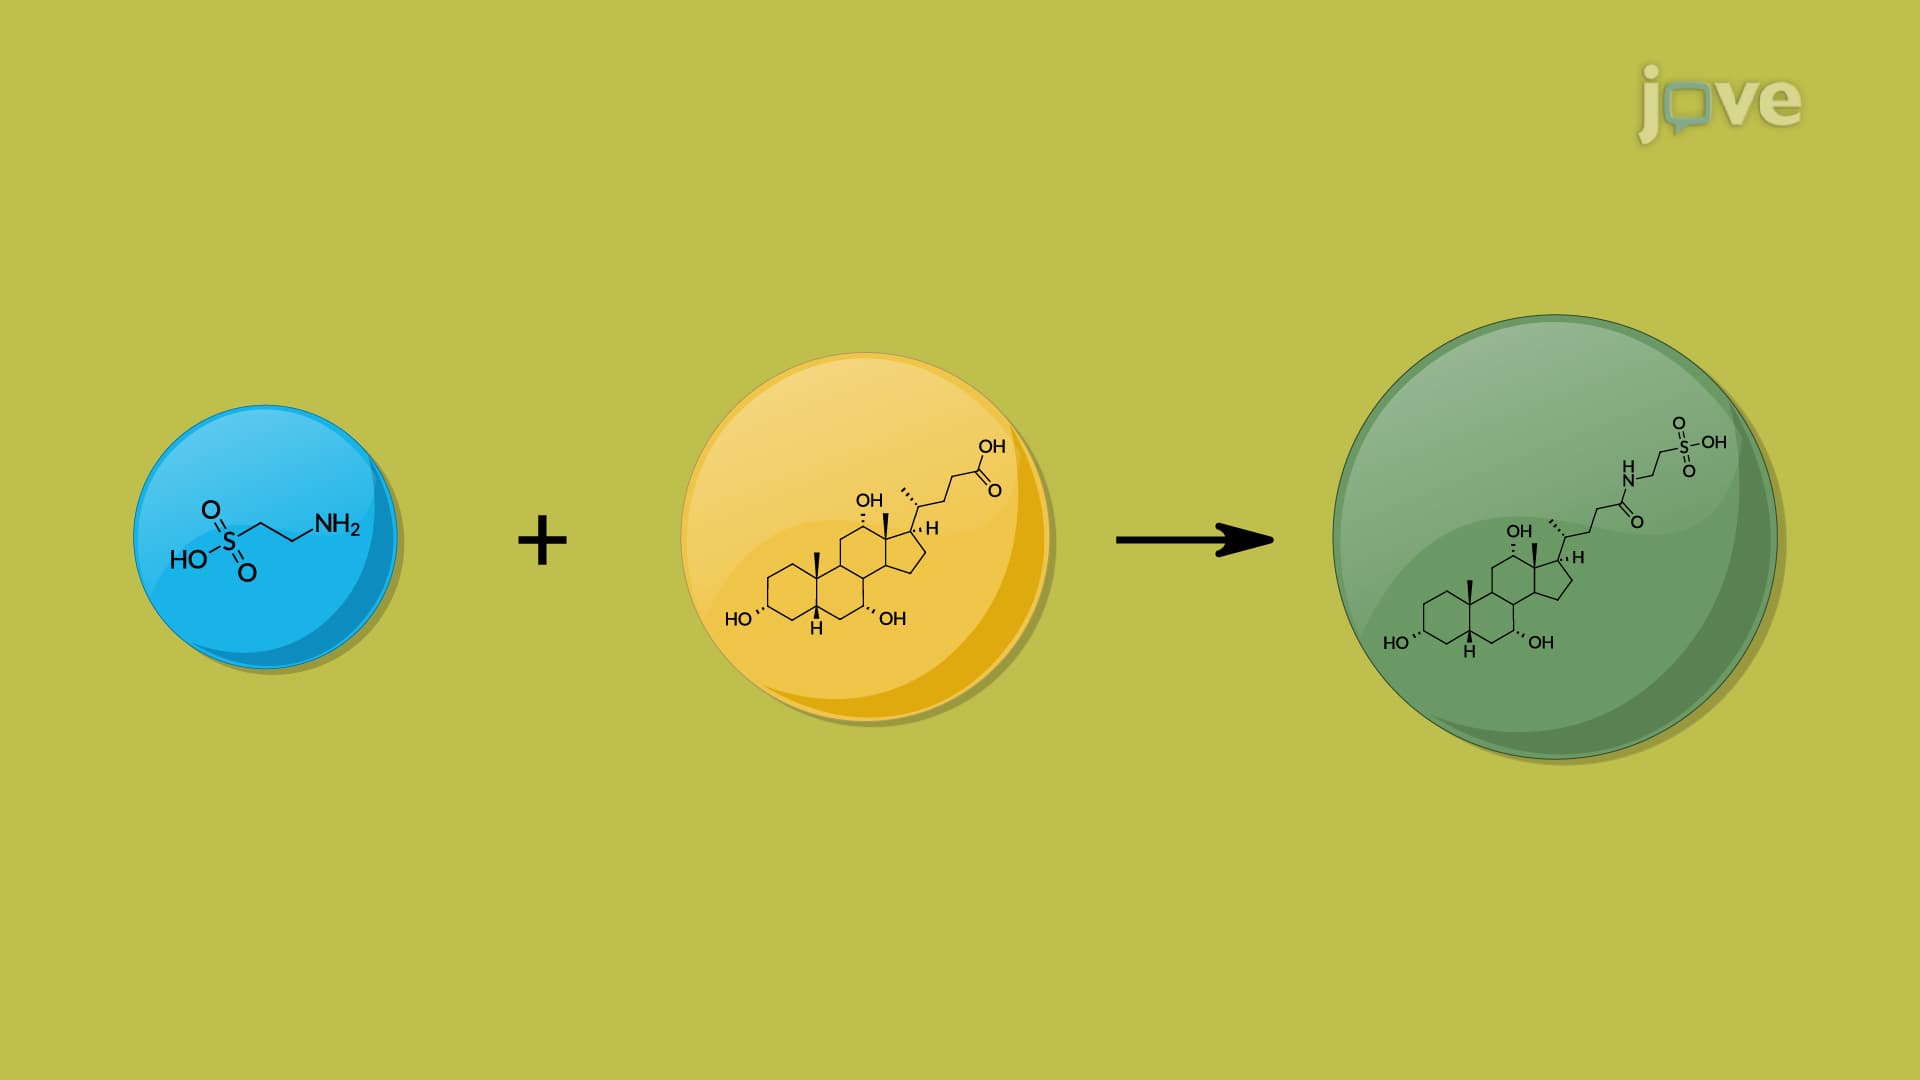
Phase II Reactions: Miscellaneous Conjugation Reactions
A key example involves the conjugation of cyanide ions, which impair cellular respiration and alter hemoglobin into non-oxygen-carrying cyanmethemoglobin. To neutralize this threat, a sulfur atom from thiosulphate is transferred to the cyanide ion, catalyzed by the enzyme rhodanese, resulting in an inactive compound called thiocyanate. The production of...
 01:28
Pharmacogenetics of Phase I Enzymes: Cytochrome P450 Isozymes
 01:27
Pharmacogenetics of Phase II Enzymes: N-acetyltransferase, Thiopurine S-methyltransferase, UDP-glucuronosyltransferase