Related Experiment Video
Updated: Aug 1, 2026
Construction and Systematical Symmetric Studies of a Series of Supramolecular Clusters with Binary or Ternary Ammonium Triphenylacetates
Published on: February 15, 2016
Ab initio many-body investigation of structure and stability of two-fold rings in silicates
A B Mukhopadhyay1, M Dolg, C Oligschleger
1Institut für Theoretische Chemie, Universität zu Köln, Greinstr.4, D-50939 Köln, Germany. a.basu@uni-koeln.de

Abstract:
In this paper we present ab initio many-body calculations on the strain energy of W silica, taken as a model system for edge-sharing tetrahedral SiO(2) systems with respect to corner-sharing ones as in alpha quartz. The mean-field results were obtained using the restricted Hartree-Fock approach, while the many-body effects were taken into account by the second-order Møller-Plesset perturbation theory and the coupled-cluster approach. Correlation contributions are found to play an important role to determine the stability of edge-sharing units. The most sophisticated method used in our calculation, i.e., the coupled-cluster approach with single and double excitations, yields a strain energy of 0.0427 a.u. per Si(2)O(4) unit with respect to alpha quartz, which is even smaller than the value obtained by a previous density functional theory calculation.
More Related Videos
Related Concept Videos
 02:22
Structures of Solids
 02:42
Ionic Crystal Structures
Most monatomic ions behave as charged spheres, and their attraction for ions of opposite charge is the same in every direction. Consequently, stable structures for ionic compounds result (1) when ions of one charge are surrounded by as many ions as possible of the opposite...
 02:02
Chair Conformation of Cyclohexane
The hydrogen atoms linked to carbons are arranged in two different axial and equatorial orientations to achieve this staggered...
 02:33
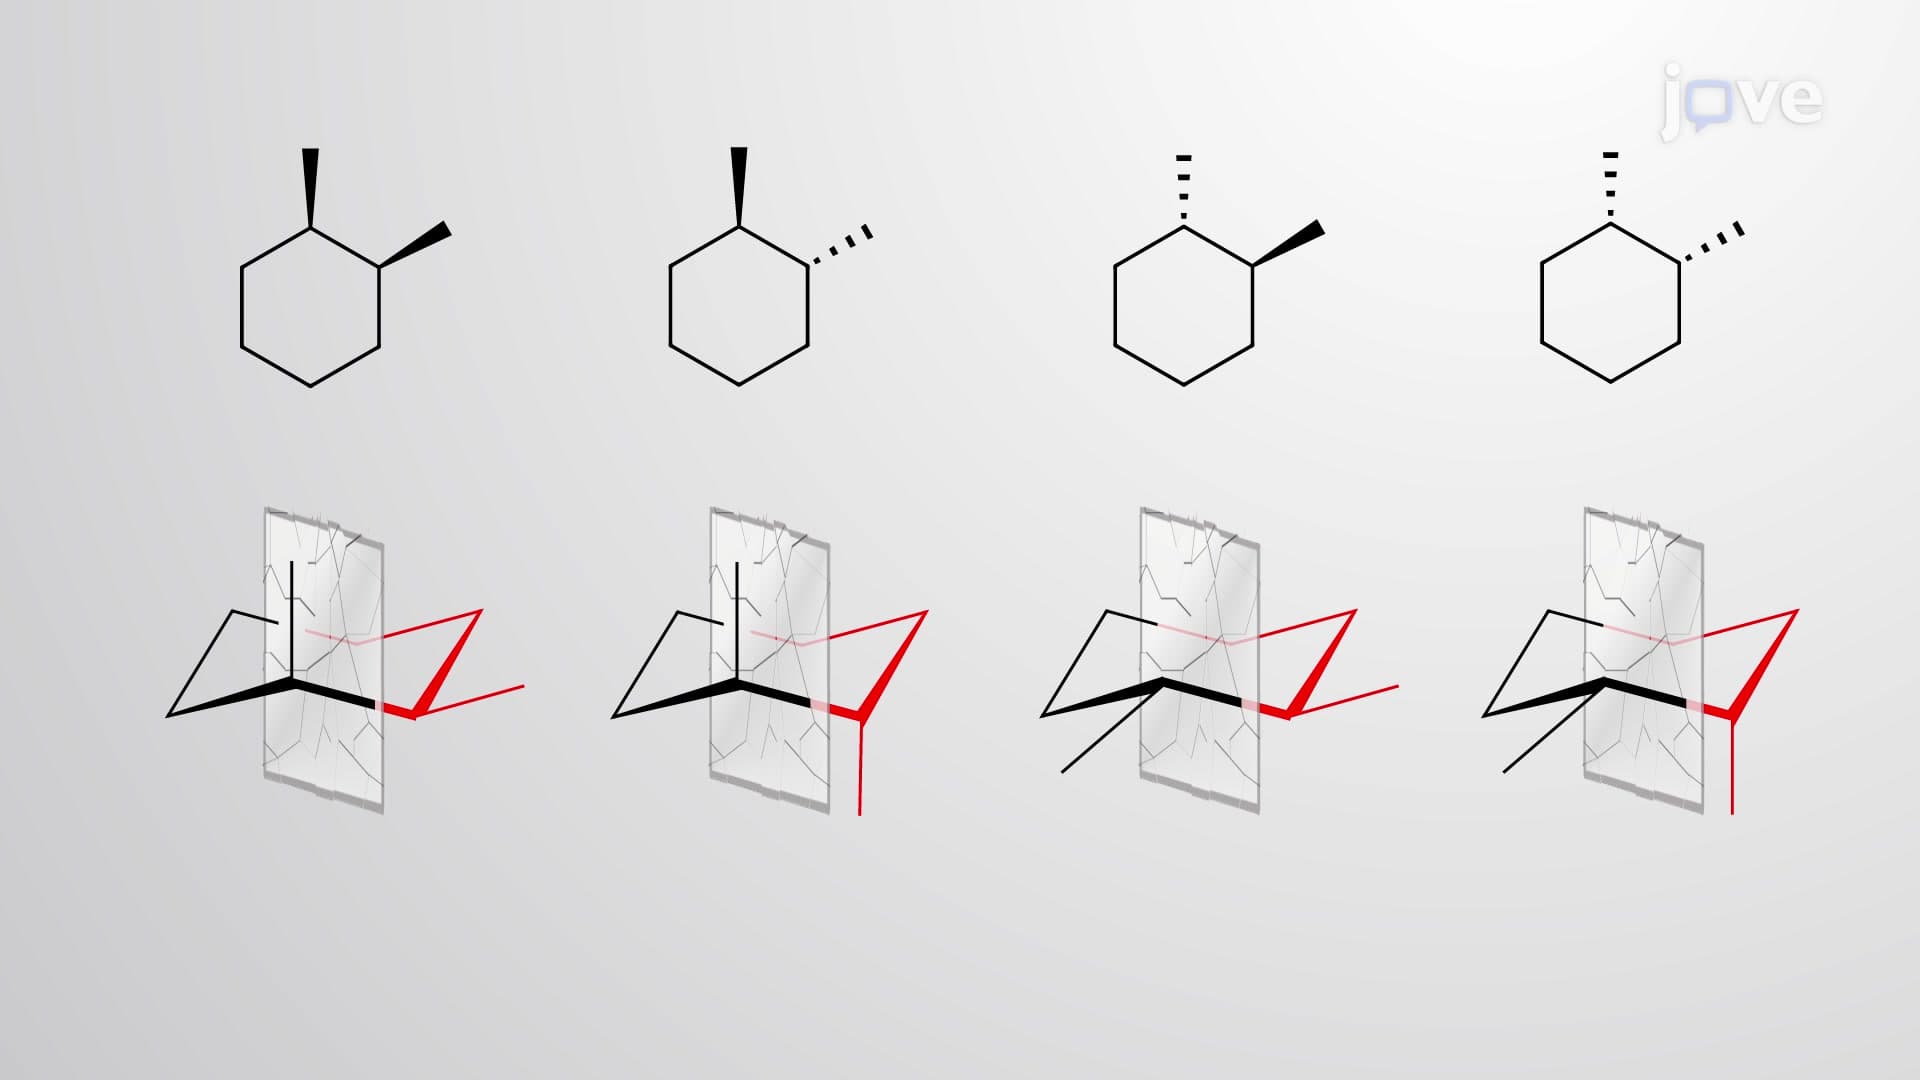
Stereoisomerism of Cyclic Compounds
 01:18
Frost Circles for Different Conjugated Systems
 01:29
Determination of Crystal Structures