Related Experiment Video
Updated: Jul 6, 2026

Investigating the Spreading and Toxicity of Prion-like Proteins Using the Metazoan Model Organism C. elegans
Published on: January 8, 2015
Prion protein disease and neuropathology of prion disease
1Neuropathology Unit, Department of Cellular Pathology and Greater Manchester Neurosciences Centre, Salford Royal Hospital, Stott Lane, Salford, M6 8HD, UK. daniel.duplessis@srft.nhs.uk

Abstract:
Human prion diseases, in common with other neurodegenerative diseases, may be sporadic or inherited and are characterized by the accumulation of cellular proteins accompanied by neuronal death and synaptic loss. Prion diseases are, however, unique in being transmissible. Central to the pathogenesis of all forms of prion disease is the prion protein. This article provides a brief overview of the biology of human prion diseases followed by a more in-depth discussion of the neuropathology of these diseases, including features of neuroradiologic relevance.
Related Concept Videos
 03:03
Amyloid Fibrils
Amyloid deposits were observed as early as 1639 in the liver and the spleen. In 1854, Rudolph Virchow performed iodine staining, normally used to...
03:03Amyloid Fibrils
Amyloid deposits were observed as early as 1639 in the liver and the spleen. In 1854, Rudolph Virchow performed iodine staining, normally used to...
 01:24
Parkinson Disease ll: Pathophysiology
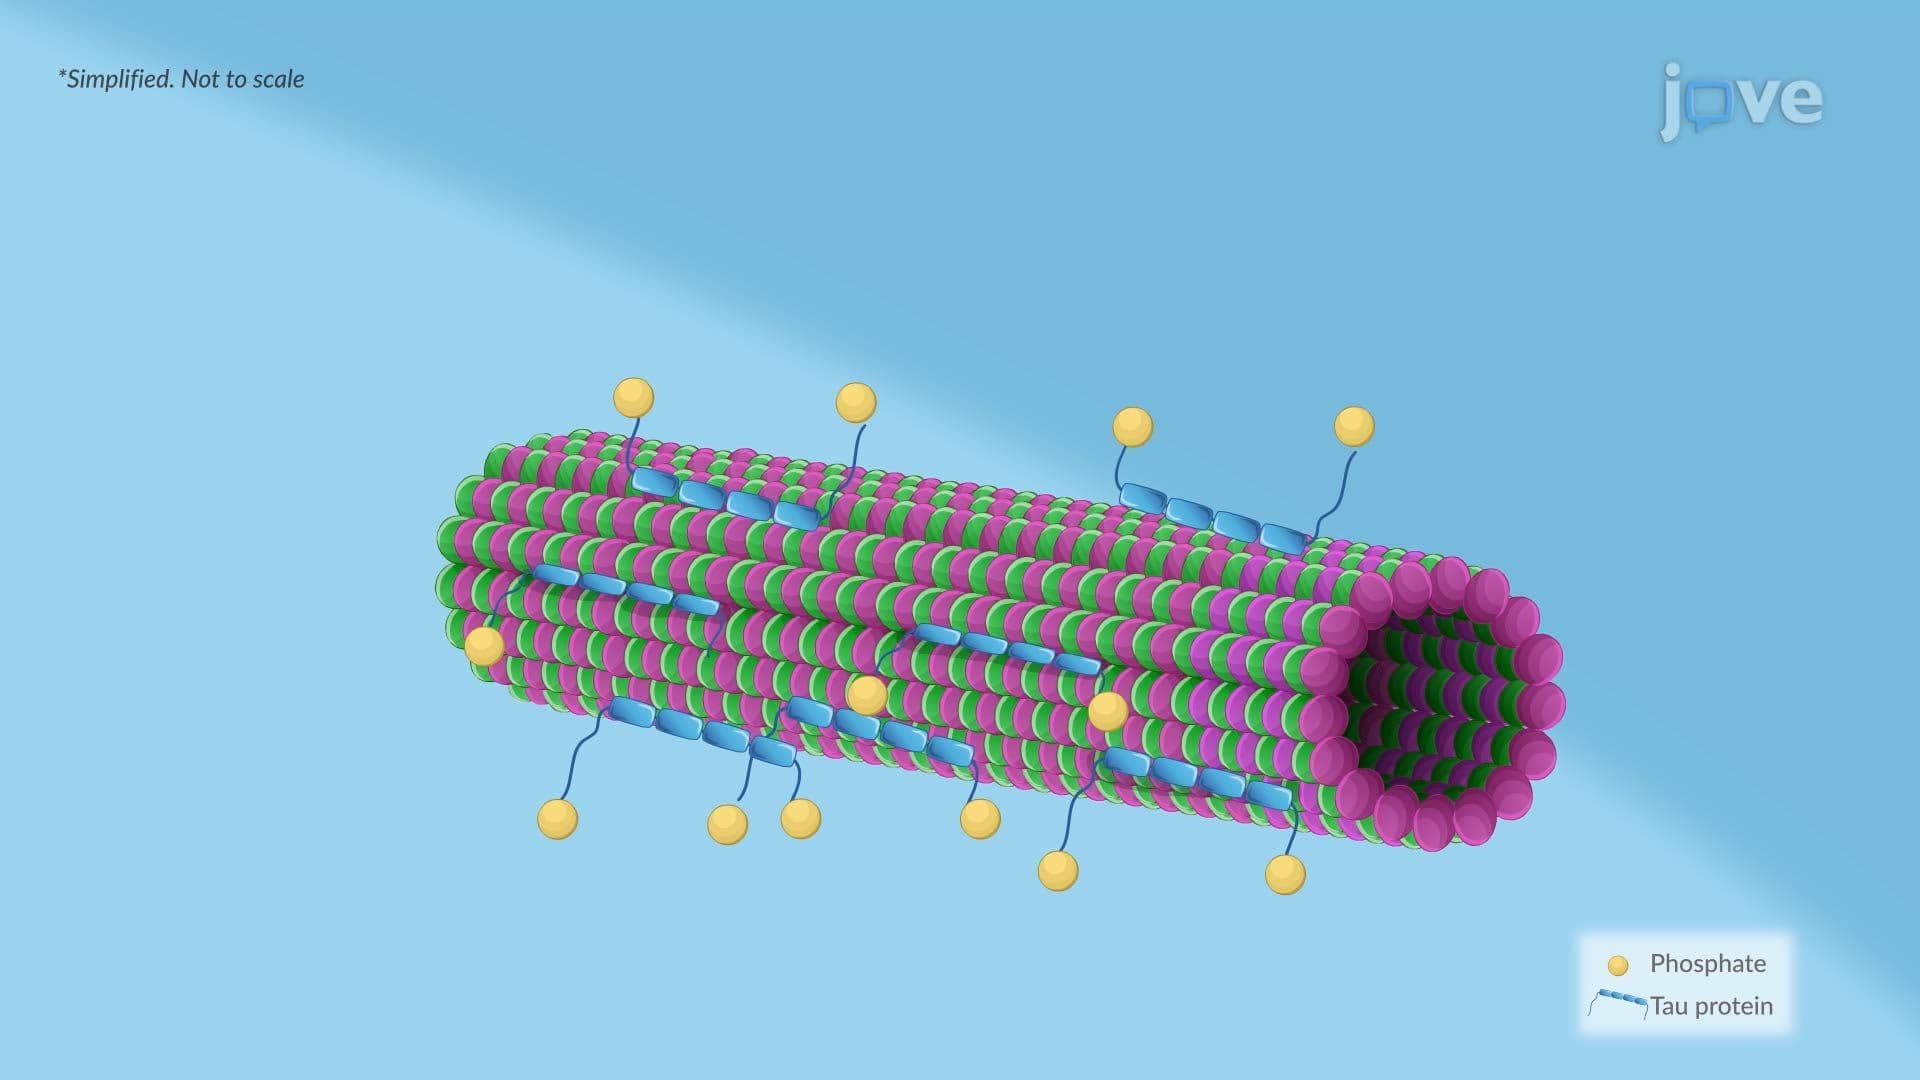 01:23
Alzheimer Disease ll: Pathophysiology
 01:29
Subviral Agents
 01:37
Neural Regulation