HemeBIND: a novel method for heme binding residue prediction by combining structural and sequence information
Related Experiment Video
Updated: Jun 1, 2026

Application of I TASSER, trRosetta, UCSF Chimera, HADDOCK server, and HEX loria for De Novo and In Silico Design of Proteins
Published on: July 8, 2025
1Department of Computer Science and Engineering, University of South Carolina, Columbia, SC 29208, USA.

Background:
Accurate prediction of binding residues involved in the interactions between proteins and small ligands is one of the major challenges in structural bioinformatics. Heme is an essential and commonly used ligand that plays critical roles in electron transfer, catalysis, signal transduction and gene expression. Although much effort has been devoted to the development of various generic algorithms for ligand binding site prediction over the last decade, no algorithm has been specifically designed to complement experimental techniques for identification of heme binding residues. Consequently, an urgent need is to develop a computational method for recognizing these important residues.
Results:
Here we introduced an efficient algorithm HemeBIND for predicting heme binding residues by integrating structural and sequence information. We systematically investigated the characteristics of binding interfaces based on a non-redundant dataset of heme-protein complexes. It was found that several sequence and structural attributes such as evolutionary conservation, solvent accessibility, depth and protrusion clearly illustrate the differences between heme binding and non-binding residues. These features can then be separately used or combined to build the structure-based classifiers using support vector machine (SVM). The results showed that the information contained in these features is largely complementary and their combination achieved the best performance. To further improve the performance, an attempt has been made to develop a post-processing procedure to reduce the number of false positives. In addition, we built a sequence-based classifier based on SVM and sequence profile as an alternative when only sequence information can be used. Finally, we employed a voting method to combine the outputs of structure-based and sequence-based classifiers, which demonstrated remarkably better performance than the individual classifier alone.
Conclusions:
HemeBIND is the first specialized algorithm used to predict binding residues in protein structures for heme ligands. Extensive experiments indicated that both the structure-based and sequence-based methods have effectively identified heme binding residues while the complementary relationship between them can result in a significant improvement in prediction performance. The value of our method is highlighted through the development of HemeBIND web server that is freely accessible at http://mleg.cse.sc.edu/hemeBIND/.
More Related Videos


Related Concept Videos
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally analyses the...
01:49Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally analyses the...
 02:21
Globular and Fibrous Proteins
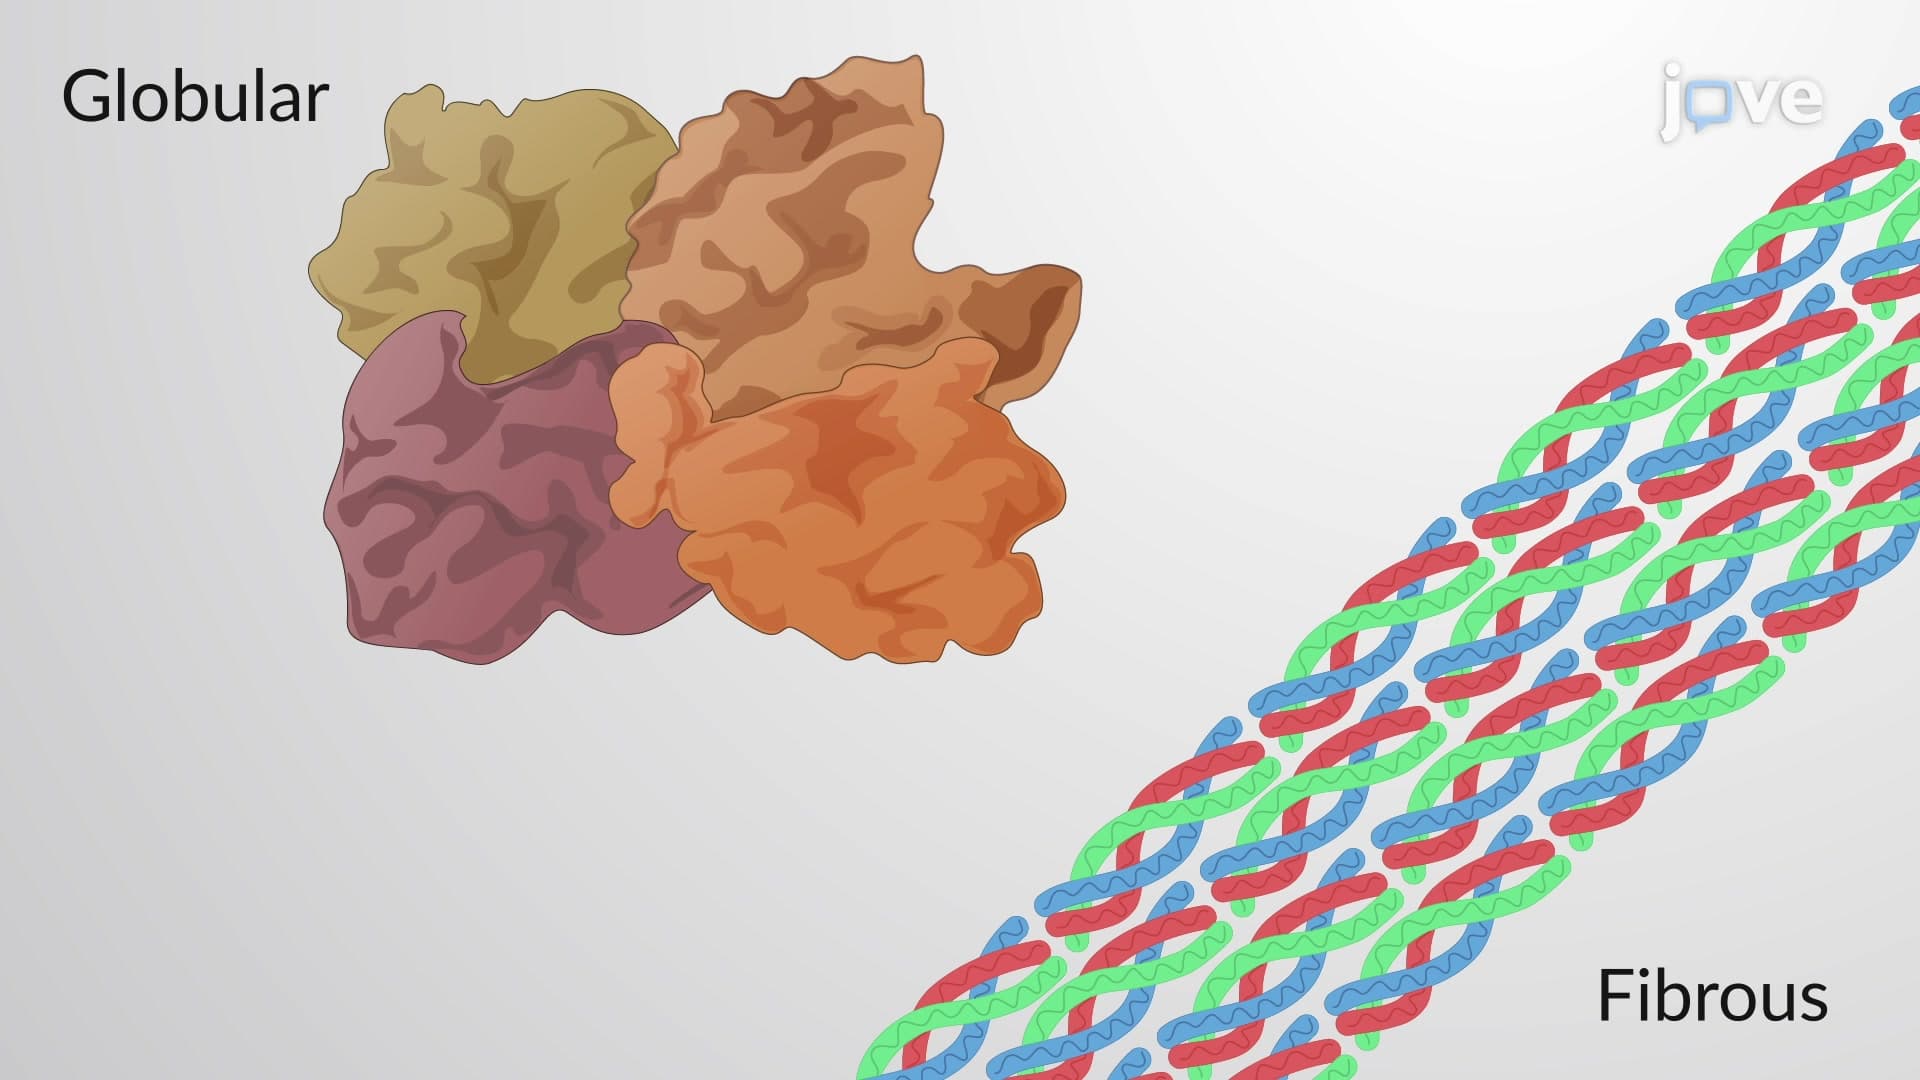
Globular proteins are also known as spheroproteins and typically are approximately round in shape. They contain a mix of amino acid types and contain differing sequences in their primary structures. Globular proteins have many different functions, such as enzymes, cellular messengers, and molecular transporters. These roles often require the proteins to be...
 01:57
Gene Families
Occasionally these regions can be adapted to take on new roles within the organism, becoming novel genes...
 02:15
Protein and Protein Structure
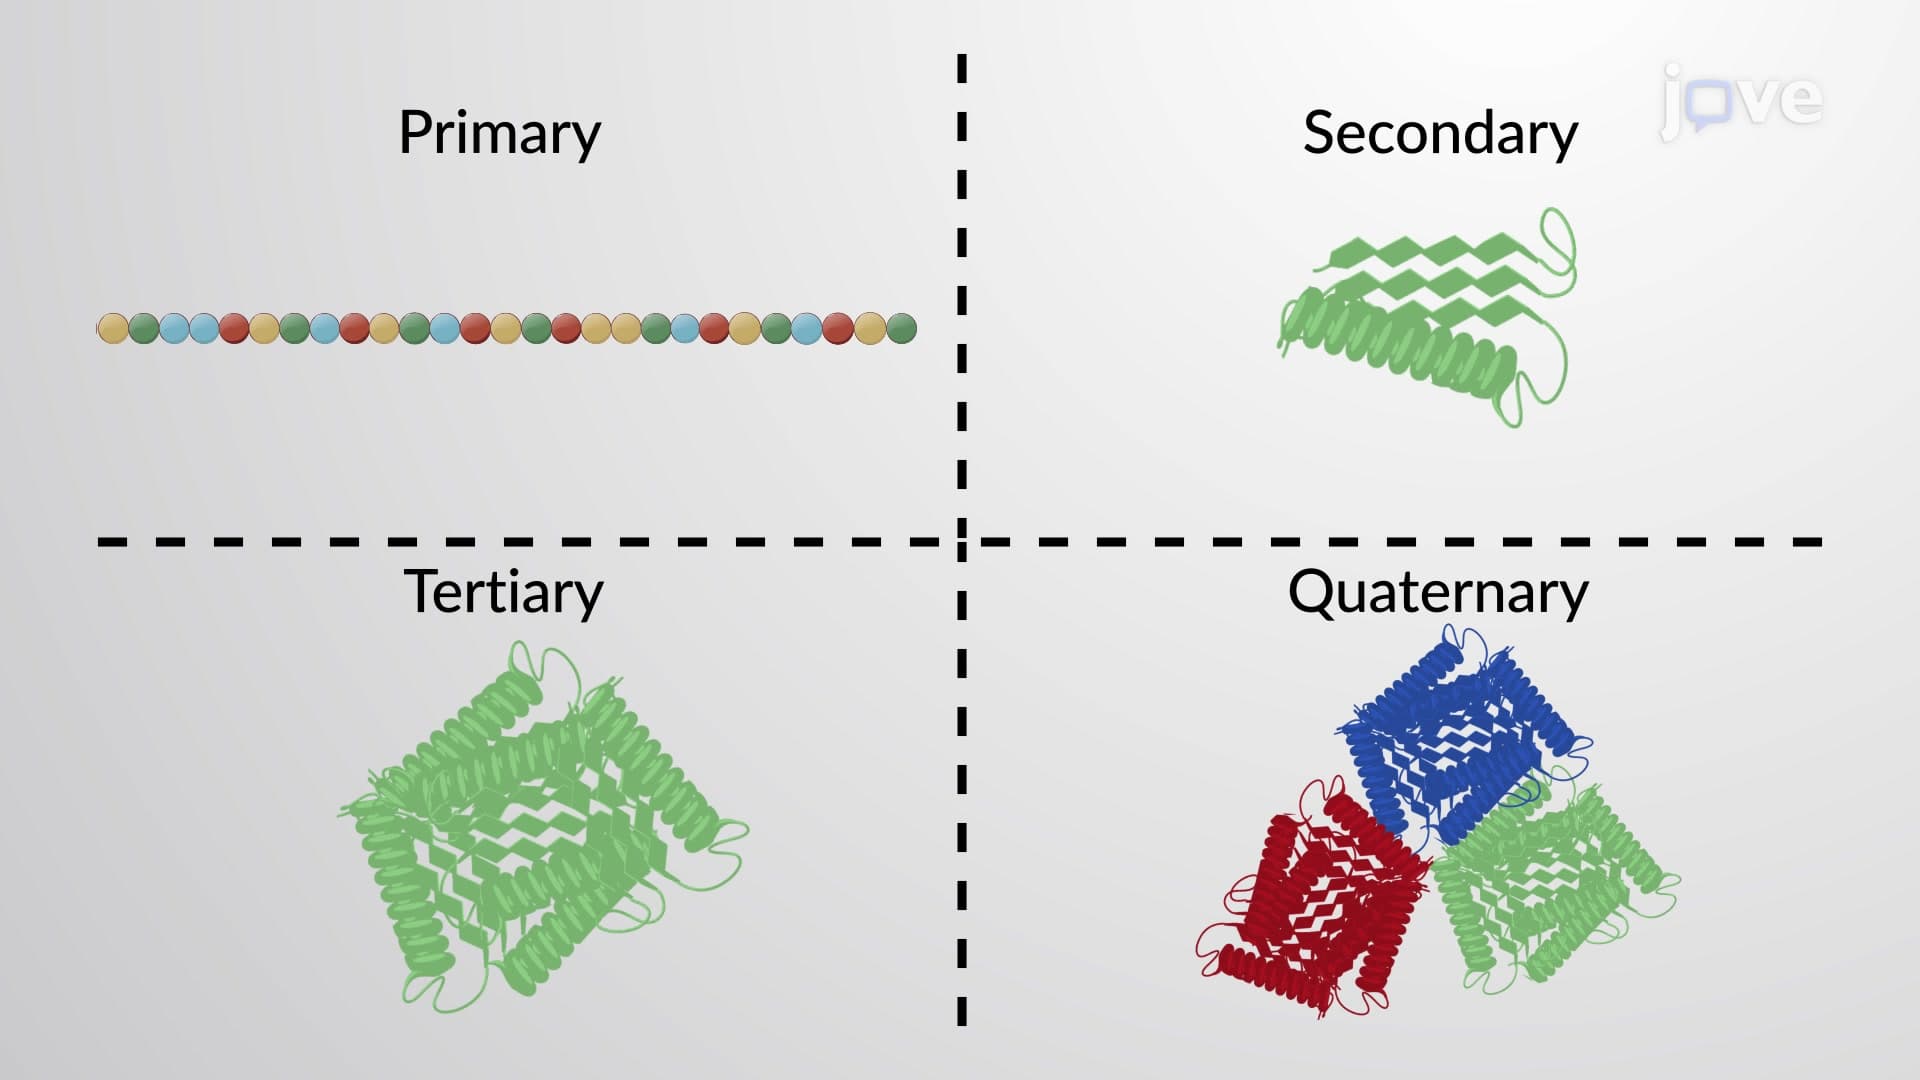
A protein's shape is critical to its function. For example, an enzyme can...
 02:40
Ligand Binding Sites
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...