Related Experiment Video
Updated: May 10, 2026

Development of Heterogeneous Enantioselective Catalysts using Chiral Metal-Organic Frameworks (MOFs)
Published on: January 17, 2020
Opposing auxiliary conformations produce the same torquoselectivity in an oxazolidinone-directed Nazarov cyclization
Bernard L Flynn1, Narasimhulu Manchala, Elizabeth H Krenske
1Medicinal Chemistry, Monash Institute of Pharmaceutical Sciences, Monash University, 381 Royal Parade, Parkville, VIC 3052, Australia. bernard.flynn@monash.edu

Abstract:
Most applications of chiral oxazolidinone auxiliaries in asymmetric synthesis operate through a common set of stereocontrol principles. That is, the oxazolidinone is made to adopt a specific, coplanar conformation with respect to the prochiral substrate, and reaction occurs preferentially at whichever stereoheterotopic face is not blocked by the substituents on the oxazolidinone. In contrast to these principles, we report here the discovery of an alternative mechanism of oxazolidinone-based stereocontrol that does not require coplanarity and is driven instead by allylic strain. This pathway has been uncovered through computational studies of an asymmetric Nazarov cyclization. Chiral oxazolidinone auxiliaries provide essentially complete control over the torquoselectivity of ring closure and the regioselectivity of subsequent deprotonation. Density functional theory calculations (M06-2X//B3LYP) reveal that in the transition state of 4π electrocyclic ring closure, the oxazolidinone ring and the cyclizing pentadienyl cation are distorted from coplanarity in a manner that gives two transition state conformations of similar energy. These two conformers are distinguished by a 180° flip in the auxiliary orientation such that in one conformer the oxazolidinone carbonyl is oriented toward the OH of the pentadienyl cation (syn-conformer) and in the other it is oriented away from this OH (anti-conformer). Surprisingly, both conformations induce the same sense of torquoselectivity, with a 3-5 kcal/mol preference for the C5-β epimer of the ring-closed cation. In both conformations, the conrotatory mode that leads to the C5-α epimer is disfavored due to higher levels of allylic strain between the oxazolidinone substituent and adjacent groups on the pentadienyl cation (R(4) and OH). The excellent torquoselectivities obtained in the oxazolidinone-directed Nazarov cyclization suggest that the allylic strain-driven stereoinduction pathway represents a viable alternative mechanism of stereocontrol for reactions of sterically congested substrates that lie outside of the traditional coplanar (N-acyloxazolidinone) paradigm.
More Related Videos
Related Concept Videos
 01:34
Woodward–Hoffmann Selection Rules and Microscopic Reversibility
 02:36
Regioselectivity and Stereochemistry of Hydroboration
Hydroboration proceeds in a concerted fashion with the attack of borane on the π bond, giving a cyclic four-centered transition state. The –BH2 group is bonded to the less substituted carbon and –H to the more substituted carbon. The concerted nature requires the simultaneous addition of –H and –BH2 across the same face of the alkene giving syn stereochemistry.
 02:23
SN2 Reaction: Stereochemistry
If the substrate is an achiral molecule at the α-carbon, the inversion of configuration is not observed.
 01:11
ortho–para-Directing Activators: –CH3, –OH, –NH2, –OCH3
 01:15
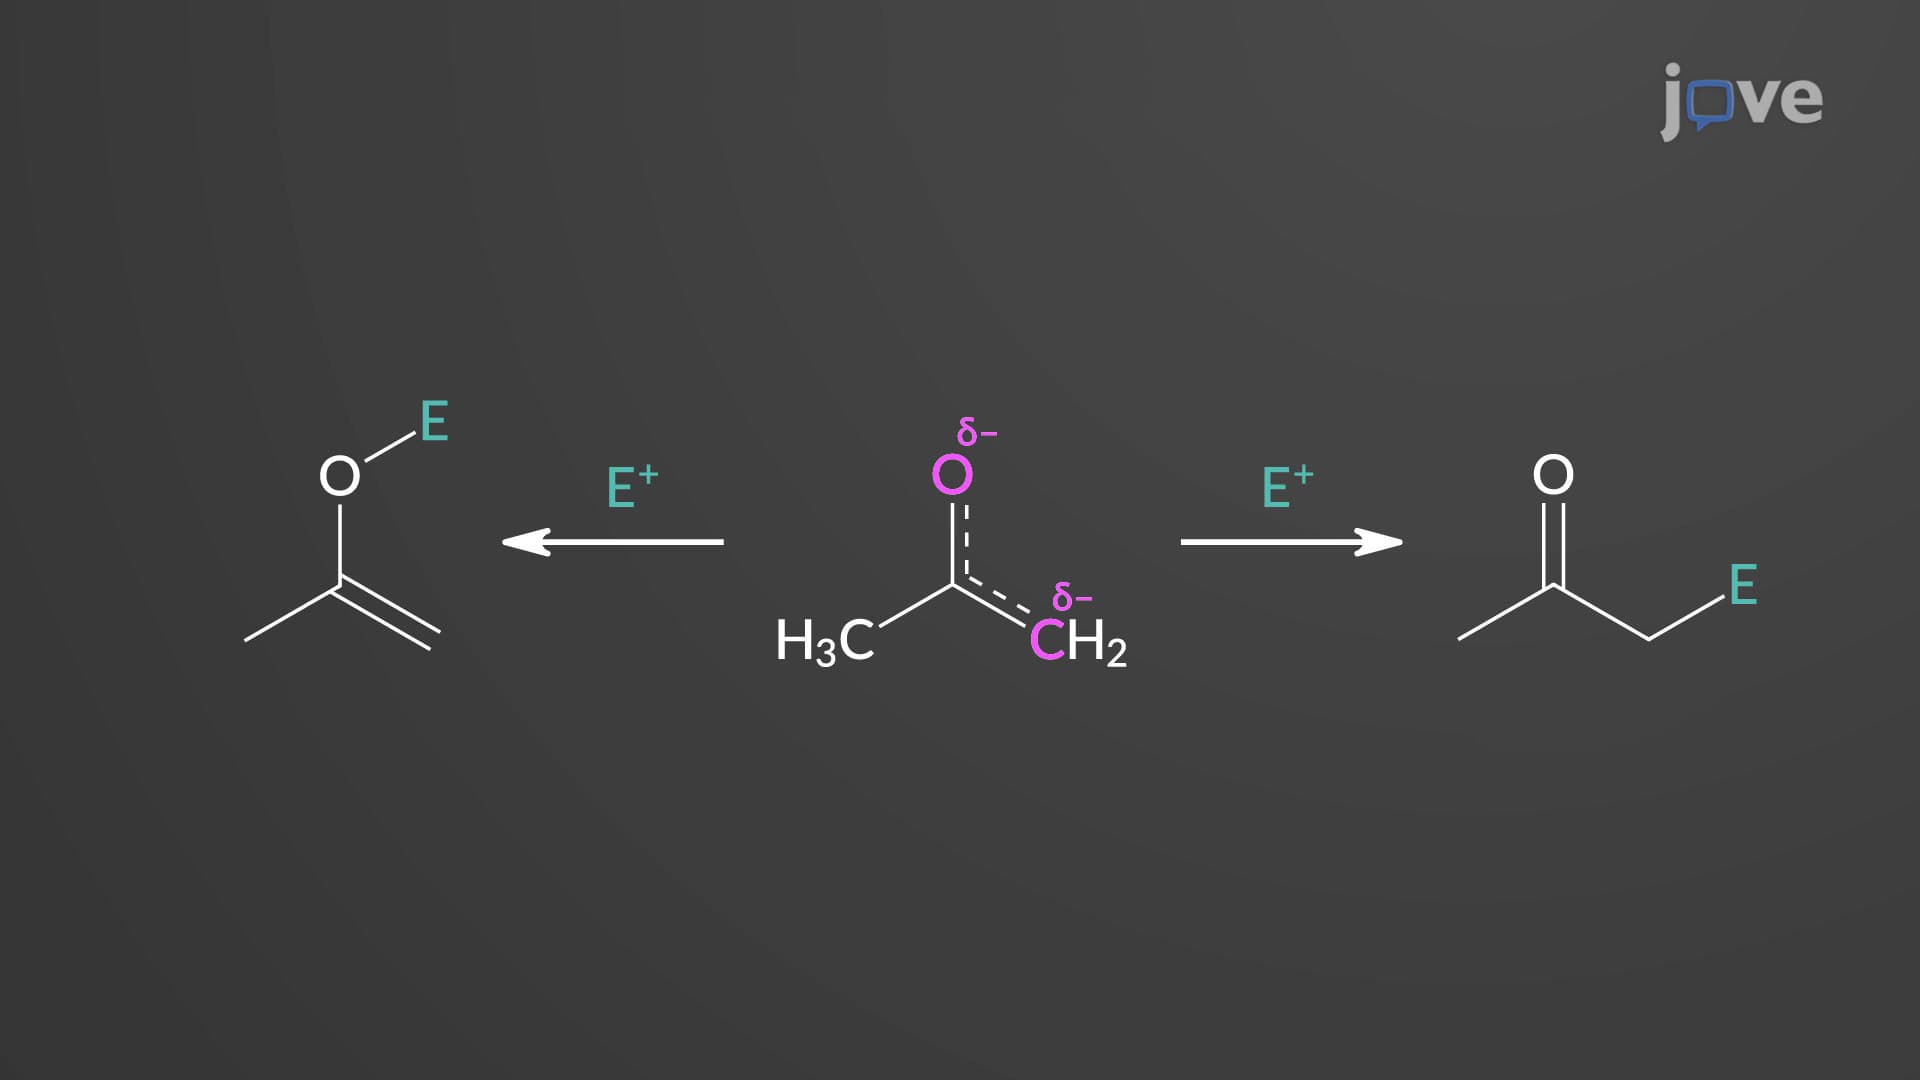
Enolate Mechanism Conventions
C-attack...
 01:17
Thermal Electrocyclic Reactions: Stereochemistry
Selection Rules: Thermal Activation
Conjugated systems containing an even number of π-electron pairs undergo a conrotatory ring closure. For example, thermal electrocyclization of (2E,4E)-2,4-hexadiene, a conjugated diene containing two π-electron pairs, gives trans-3,4-dimethylcyclobutene.