Related Experiment Video
Updated: Nov 17, 2025
Author Spotlight: Streamlining Protein Target Prediction and Validation via Molecular Docking and CETSA
Published on: February 23, 2024
Predicting Drug-Target Affinity Based on Recurrent Neural Networks and Graph Convolutional Neural Networks
Qingyu Tian1, Mao Ding2, Hui Yang3
1College of Computer Science and Technology, China University of Petroleum, Qingdao 266580, Shandong,China.

Background:
Drug development requires a lot of money and time, and the outcome of the challenge is unknown. So, there is an urgent need for researchers to find a new approach that can reduce costs. Therefore, the identification of drug-target interactions (DTIs) has been a critical step in the early stages of drug discovery. These computational methods aim to narrow the search space for novel DTIs and to elucidate the functional background of drugs. Most of the methods developed so far use binary classification to predict the presence or absence of interactions between the drug and the target. However, it is more informative but also more challenging to predict the strength of the binding between a drug and its target. If the strength is not strong enough, such a DTI may not be useful. Hence, the development of methods to predict drug-target affinity (DTA) is of significant importance Method: We have improved the GraphDTA model from a dual-channel model to a triple-channel model. We interpreted the target/protein sequences as time series and extracted their features using the LSTM network. For the drug, we considered both the molecular structure and the local chemical background, retaining the four variant networks used in GraphDTA to extract the topological features of the drug and capturing the local chemical background of the atoms in the drug by using BiGRU. Thus, we obtained the latent features of the target and two latent features of the drug. The connection of these three feature vectors is then inputted into a 2 layer FC network, and a valuable binding affinity is the output.
Result:
We used the Davis and Kiba datasets, using 80% of the data for training and 20% of the data for validation. Our model showed better performance when compared with the experimental results of GraphDTA Conclusion: In this paper, we altered the GraphDTA model to predict drug-target affinity. It represents the drug as a graph and extracts the two-dimensional drug information using a graph convolutional neural network. Simultaneously, the drug and protein targets are represented as a word vector, and the convolutional neural network is used to extract the time-series information of the drug and the target. We demonstrate that our improved method has better performance than the original method. In particular, our model has better performance in the evaluation of benchmark databases.
More Related Videos
 05:47
Evidence-based Knowledge Synthesis and Hypothesis Validation: Navigating Biomedical Knowledge Bases via Explainable AI and Agentic Systems
Published on: June 13, 2025
 07:15
Machine Learning Algorithms for Early Detection of Bone Metastases in an Experimental Rat Model
Published on: August 16, 2020
Related Concept Videos
 02:26
Protein Networks
These interactions can be represented through maps depicting protein-protein interaction networks, represented as nodes and edges. Nodes are circles that are representative of a protein,...
 01:29
Drug-Receptor Interactions
Several parameters, such as the drug's affinity for its receptor and its efficacy, which is its ability to activate the receptor, determine the drug's effect on the tissue....
 01:26
Drug Discovery: Overview
 01:26
Targets for Drug Action: Overview
Receptors are either membrane-spanning or intracellular proteins, which upon binding a ligand, get activated and transmit the signal downstream to elicit a response. Drugs bind receptors, either mimicking the action of endogenous ligands or blocking the receptor activity to bring about a modified response. Nearly 35% of approved drugs target the G...
 01:25
Drug-Receptor Bonds
In...
 01:25
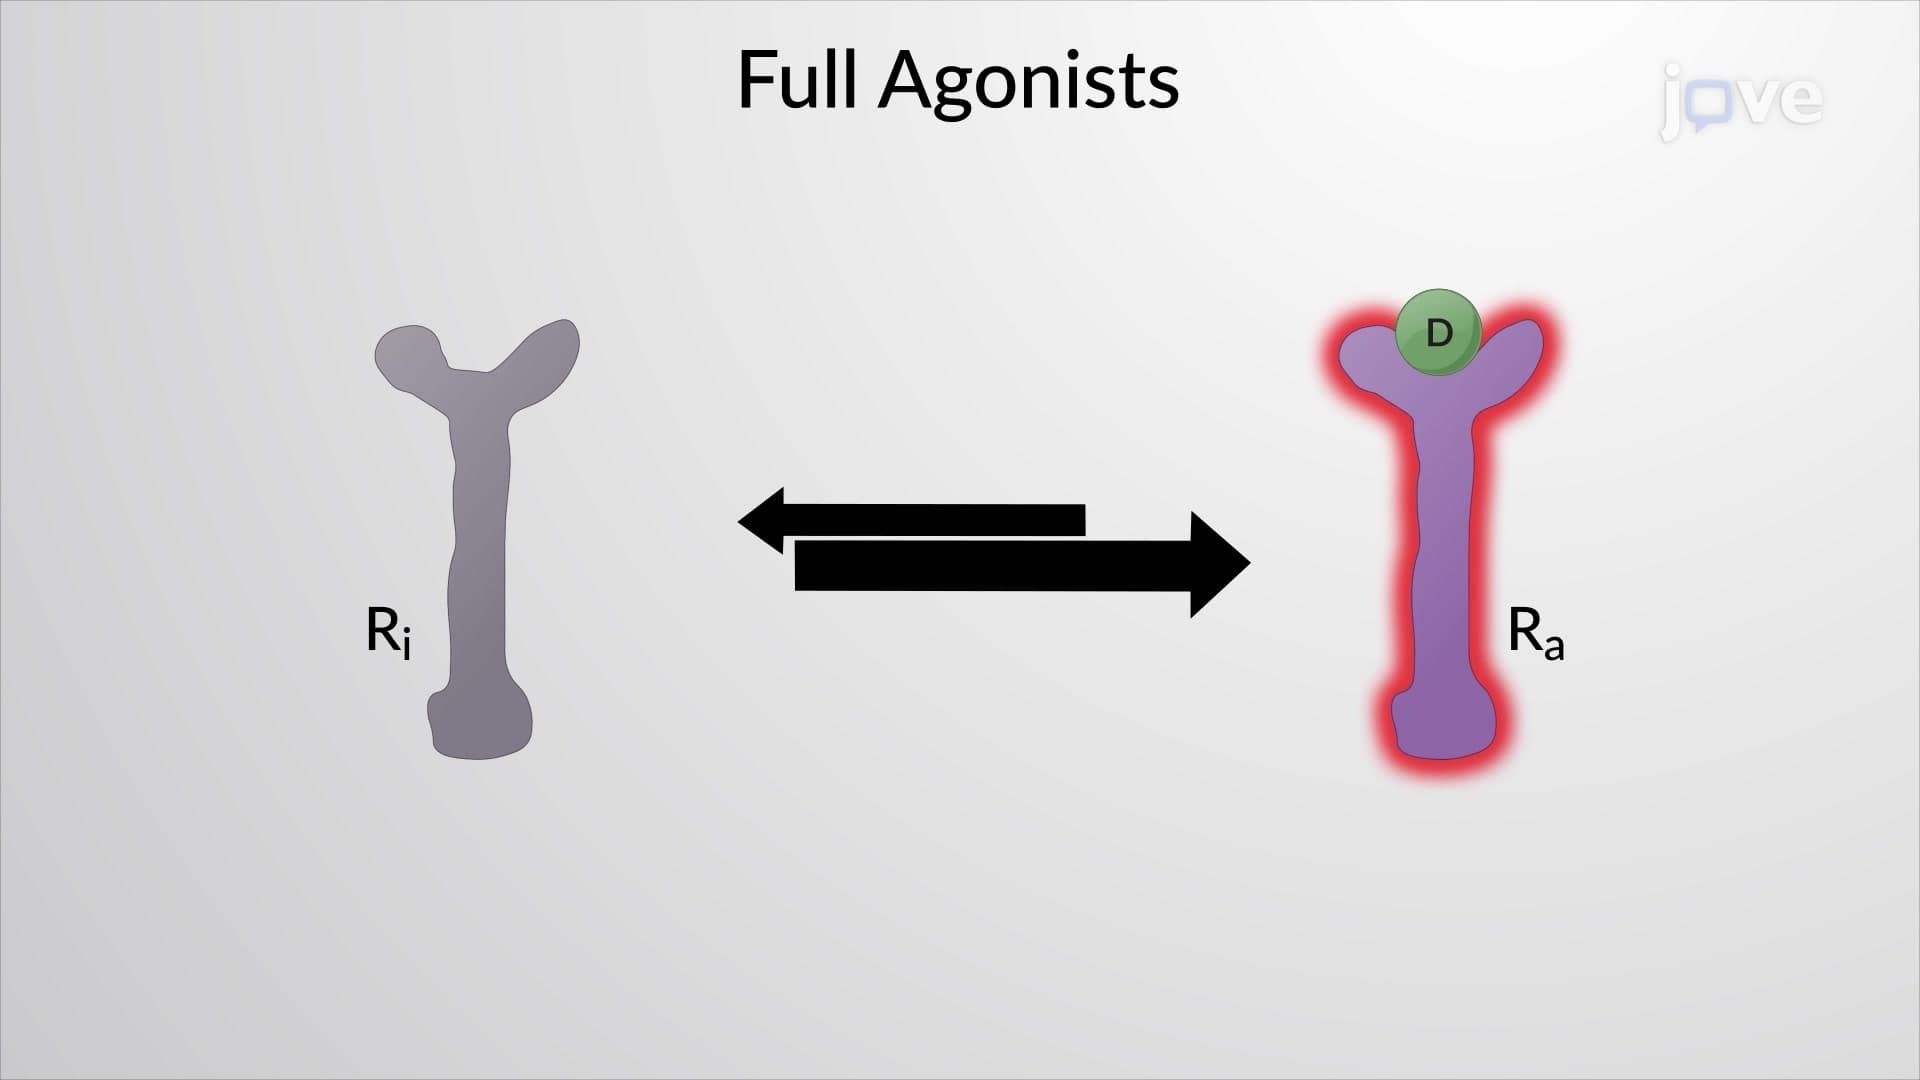
Drug-Receptor Interaction: Agonist
Agonists can bind to receptors in different ways. Some agonists bind directly to the receptor's active site, mimicking the endogenous...