Related Experiment Video
Updated: Jul 17, 2025

Probe Type II Band Alignment in One-Dimensional Van Der Waals Heterostructures Using First-Principles Calculations
Published on: October 12, 2019
The FP-LAPW/GAM-MPW1K approach: a reliable abinitio method for calculating the band gap of II-VI semiconductors
Amor Toumiat1, Abdelghani May2
1Physics Department, Constantine 1-Freres Mentouri University, Constantine, Algeria. amor.toumiat@umc.edu.dz.

Context:
The bandgap of metal monochalcogenides (MMCs) is a key property that governs their physical and chemical properties. Accurate measurement of the bandgap is essential for a range of applications, including optoelectronics and photovoltaics. However, many theoretical approximations fail to accurately calculate the bandgap for MMCs, making it difficult to obtain precise values. This study investigated the suitability of the FP-LAPW/GAM-MPW1K scheme for determining the bandgap of MMCs. The investigation included lattice parameters, bandgap, band structure, and density of states, which were compared against both previous theoretical calculations and available experimental data. The findings of the study indicate that the FP-LAPW/GAM-MPW1K approach accurately calculates the bandgap value of MMCs by efficiently treating d-state electrons. The results are consistent with prior studies, confirming the method's reliability in determining the bandgap of these semiconductors.
Methods:
our study used the GAM-MPW1K functional and the full potential linearized augmented plane wave method (FP-LAPW) in the ELK code to calculate the lattice parameters, electronic band structure, and bandgap of ZnS, ZnSe, ZnTe, CdS, CdSe, and CdTe compounds in the wurtzite structure. The crystallographic data were obtained from the COD database and the inputs were prepared by CIF2CELL code. The results were visualized using Xmgrace and VESTA software.
Related Concept Videos
 01:12
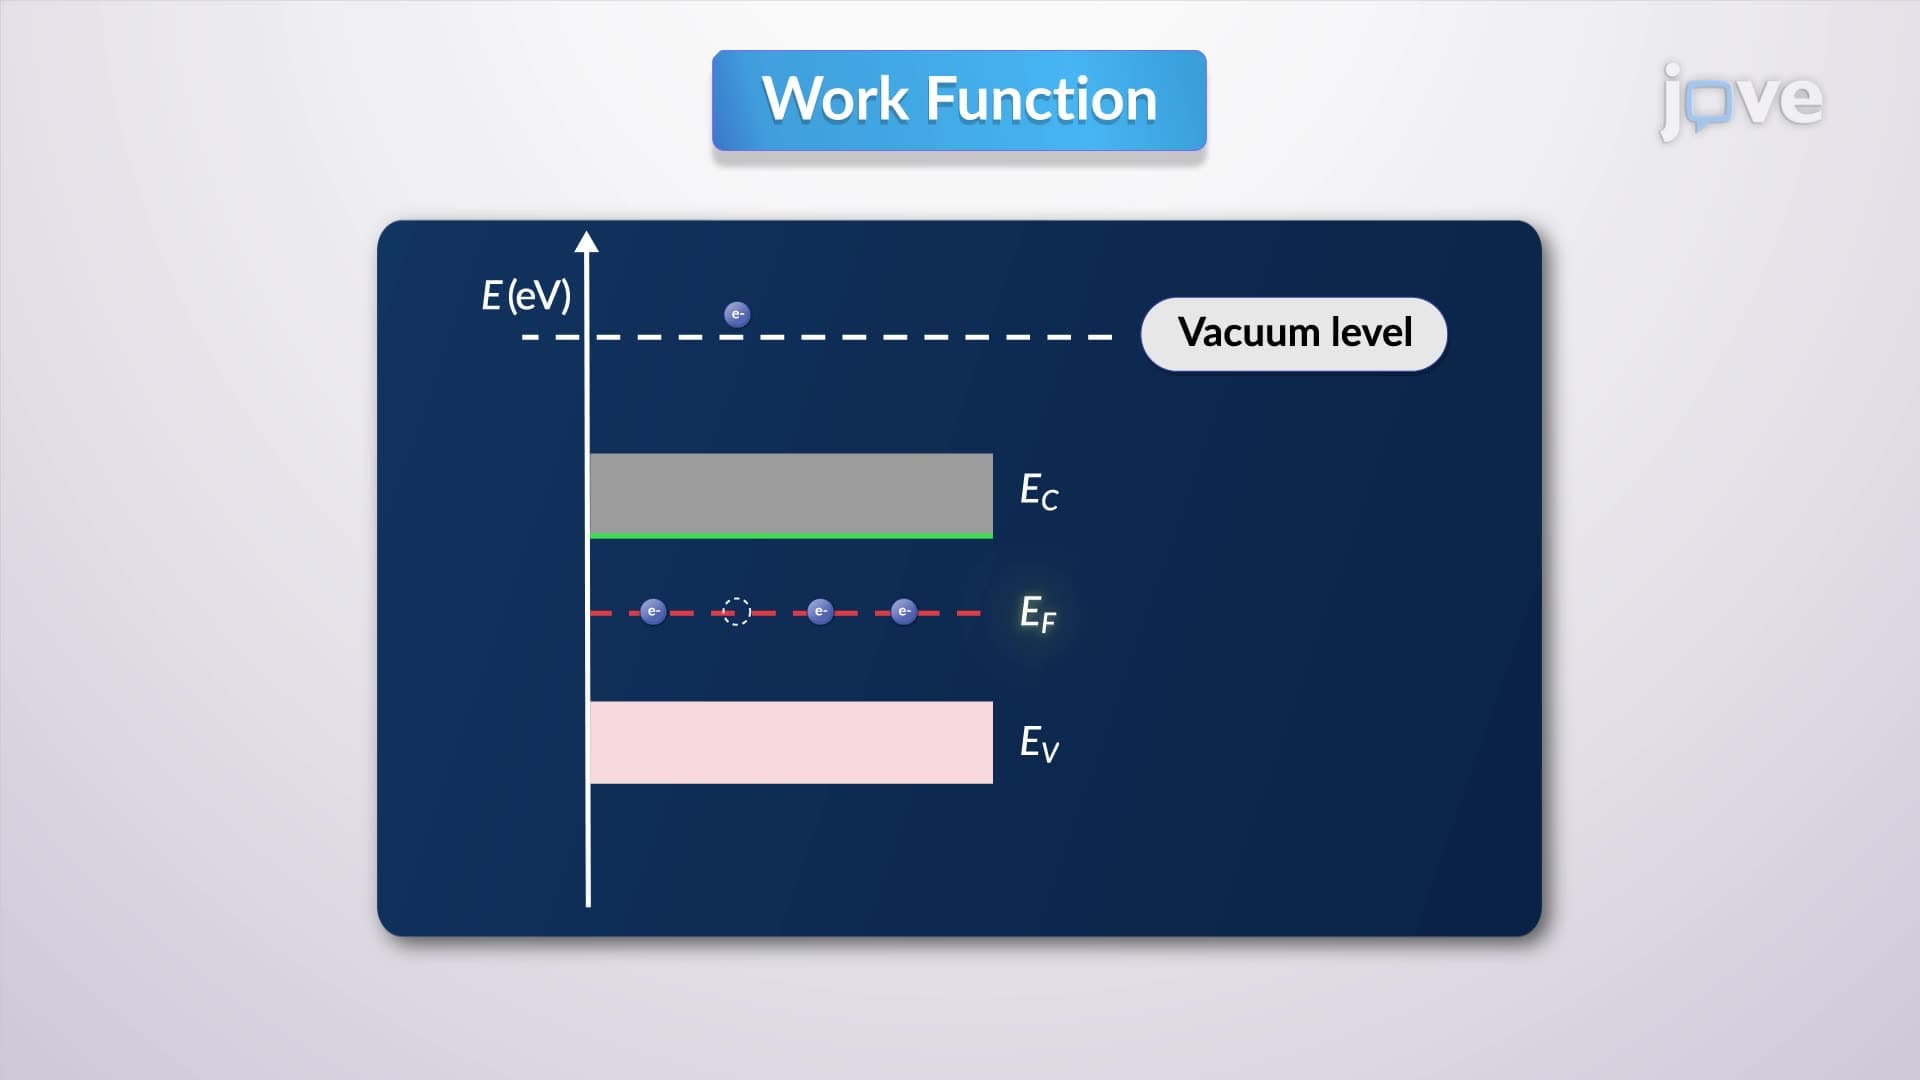
Fermi Level Dynamics
Electron affinity in semiconductors refers to the energy gap between the minimum of its conduction band and the vacuum level and it is a critical parameter in determining how easily a semiconductor can accept additional electrons.
The work...
 01:22
Semiconductors
Metals such as copper (Cu), zinc (Zn), or lead (Pb) have low resistivity and feature conduction bands that are either not fully occupied or overlap with the valence band, making a bandgap non-existent. This allows electrons in the highest energy levels of the valence band to easily transition to the conduction band upon gaining...
 02:35
Band Theory
The energy difference between these bands is known as the band gap.
Conductor, Semiconductor,...
 01:18
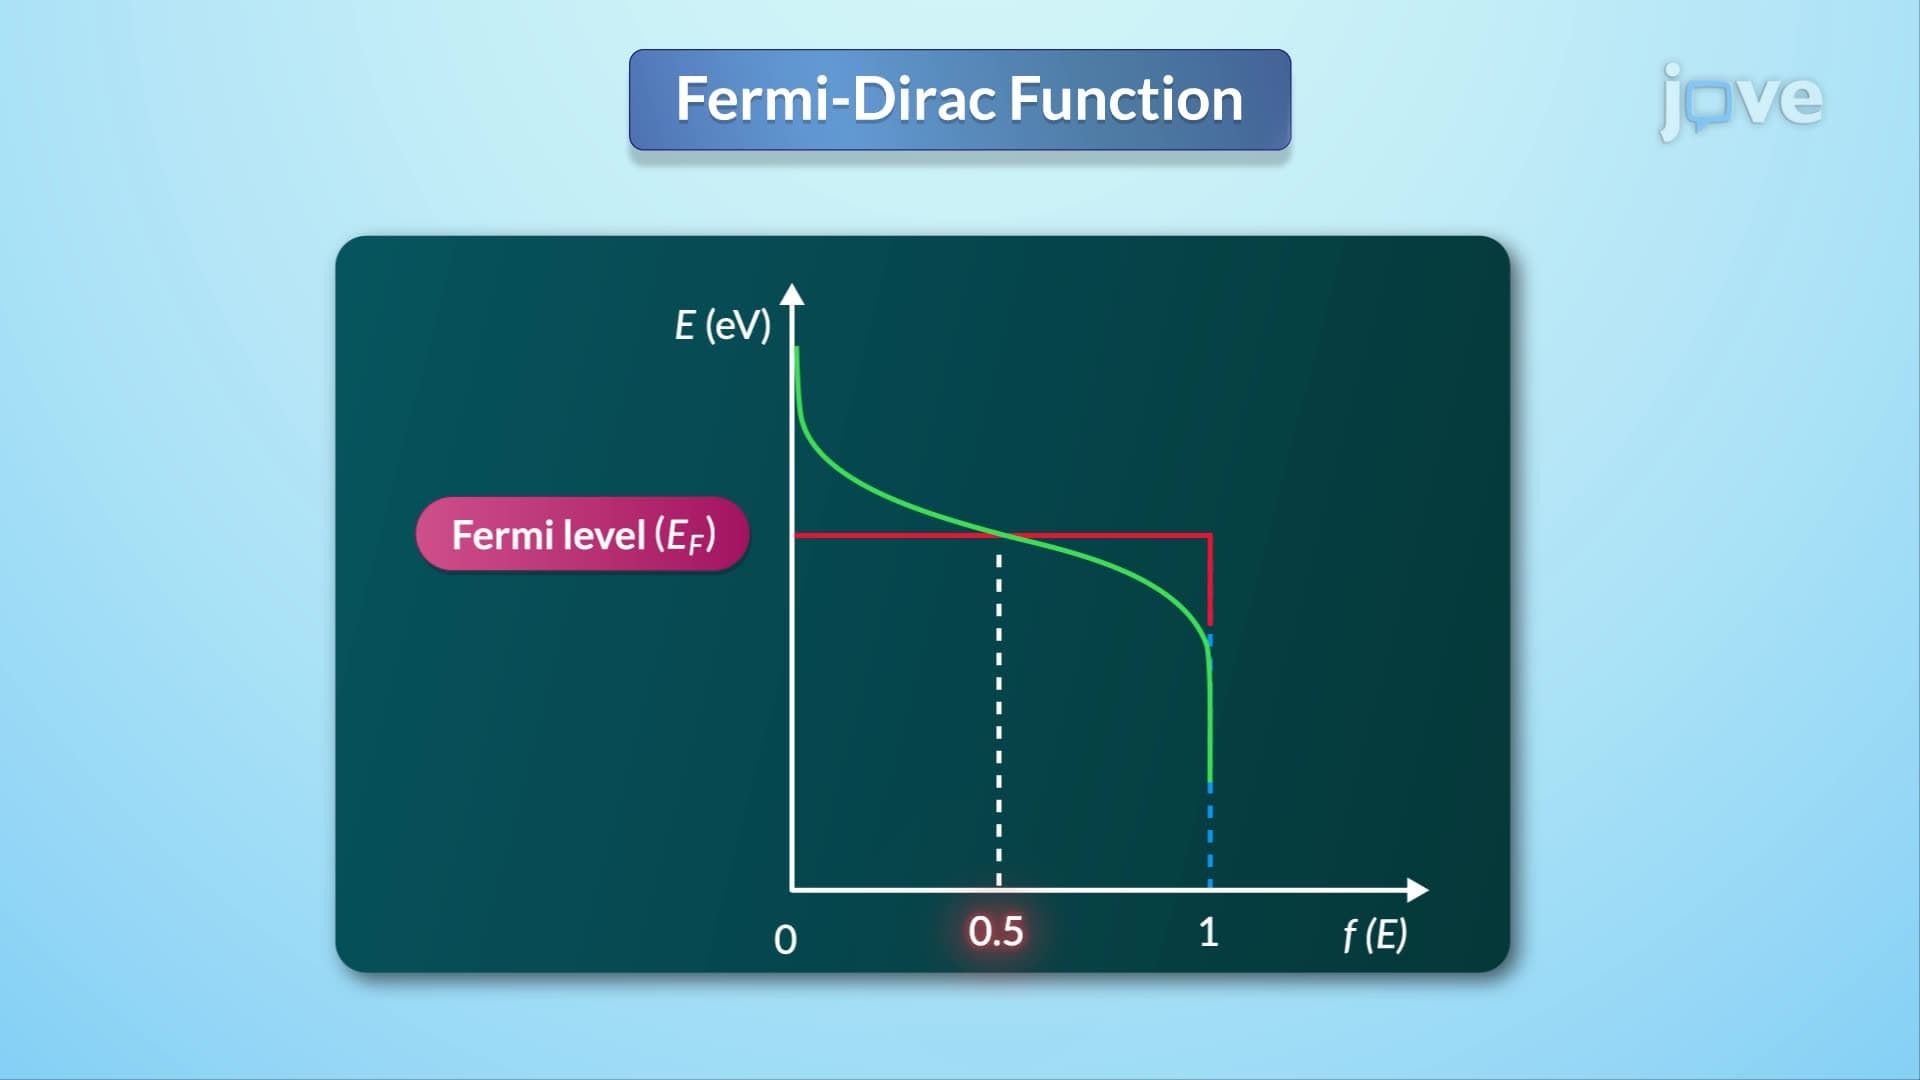
Fermi Level
At absolute zero temperature, electrons fill all energy states up to the Fermi level, leaving upper states empty. As the temperature rises,...
 01:01
Energy Bands in Solids
Band Formation:
When atoms are brought close together, as in a solid, these discrete energy levels begin to split due to the overlap of electron orbitals from adjacent atoms. This split occurs because of the Pauli exclusion principle, which states...