

Flexible protein-protein docking with a multitrack iterative transformer
Lee-Shin Chu1, Jeffrey A Ruffolo2, Ameya Harmalkar1
1Department of Chemical and Biomolecular Engineering, Johns Hopkins University, Baltimore, Maryland, USA.

Abstract:
Conventional protein-protein docking algorithms usually rely on heavy candidate sampling and reranking, but these steps are time-consuming and hinder applications that require high-throughput complex structure prediction, for example, structure-based virtual screening. Existing deep learning methods for protein-protein docking, despite being much faster, suffer from low docking success rates. In addition, they simplify the problem to assume no conformational changes within any protein upon binding (rigid docking). This assumption precludes applications when binding-induced conformational changes play a role, such as allosteric inhibition or docking from uncertain unbound model structures. To address these limitations, we present GeoDock, a multitrack iterative transformer network to predict a docked structure from separate docking partners. Unlike deep learning models for protein structure prediction that input multiple sequence alignments, GeoDock inputs just the sequences and structures of the docking partners, which suits the tasks when the individual structures are given. GeoDock is flexible at the protein residue level, allowing the prediction of conformational changes upon binding. On the Database of Interacting Protein Structures (DIPS) test set, GeoDock achieves a 43% top-1 success rate, outperforming all other tested methods. However, in the standard DIPS train/test splits, we discovered contamination of close homologs in the training set. After decontaminating the training set, the success rate is 31%. On the DB5.5 test set and a benchmark dataset of antibody-antigen complexes, GeoDock outperforms the deep learning models trained using the same dataset but falls behind most of the conventional methods and AlphaFold-Multimer. GeoDock attains an average inference speed of under 1 s on a single GPU, enabling its application in large-scale structure screening. Although binding-induced conformational changes are still a challenge owing to limited training and evaluation data, our architecture sets up the foundation to capture this backbone flexibility. Code and a demonstration Jupyter notebook are available at https://github.com/Graylab/GeoDock.
More Related Videos
 10:58
SNARE-mediated Fusion of Single Proteoliposomes with Tethered Supported Bilayers in a Microfluidic Flow Cell Monitored by Polarized TIRF Microscopy
Published on: August 24, 2016
 10:21
Author Spotlight: Streamlining Protein Target Prediction and Validation via Molecular Docking and CETSA
Published on: February 23, 2024
Related Concept Videos
 01:28
Protein Translocation Machinery on the ER Membrane
Sec61 protein conducting channel
In eukaryotes, the translocon complex comprises a core heterotrimeric translocator channel called the Sec61 complex. This channel includes three transmembrane proteins, Sec61α, Sec61β, and Sec61γ, and is the largest subunit of the...
 02:04
Protein-protein Interfaces
 01:09
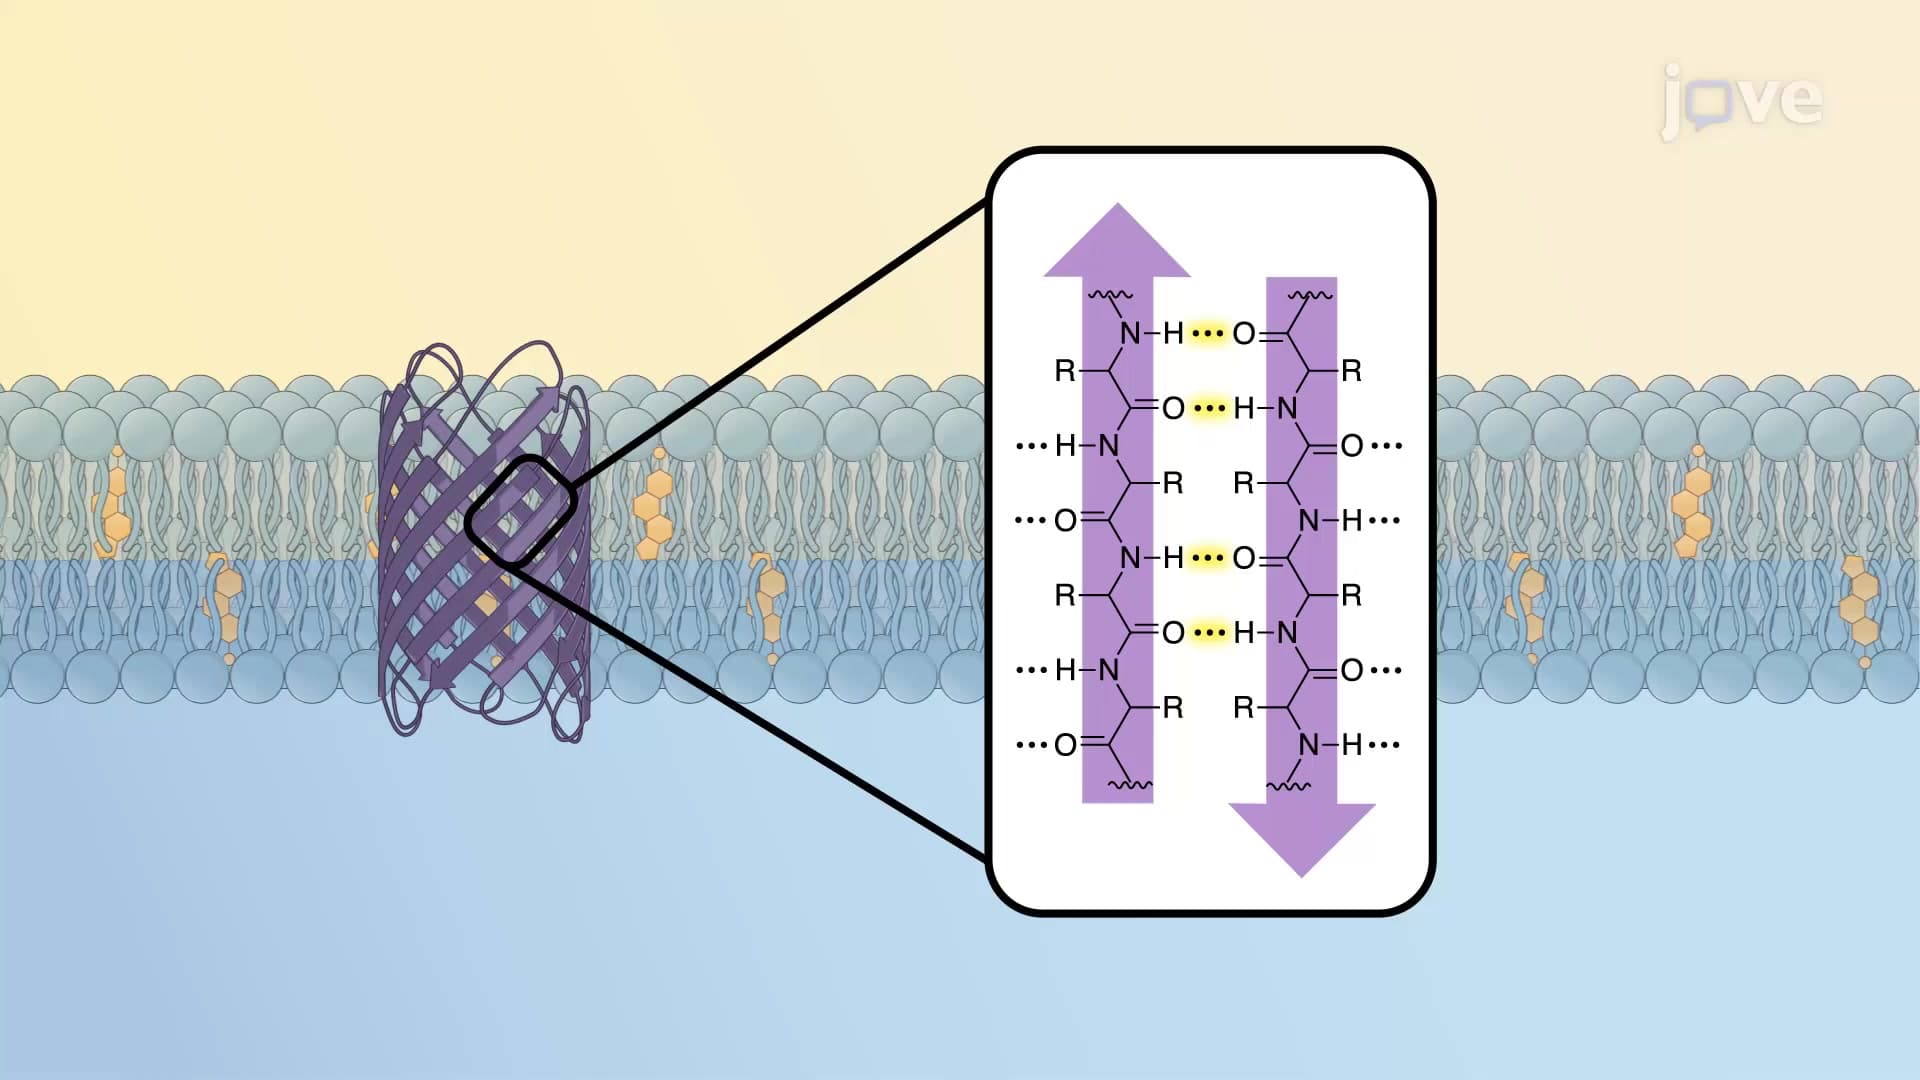
Multi-pass Transmembrane Proteins and β-barrels
α-Helix containing multi-pass transmembrane proteins
Multi-pass transmembrane proteins such as...
 01:29
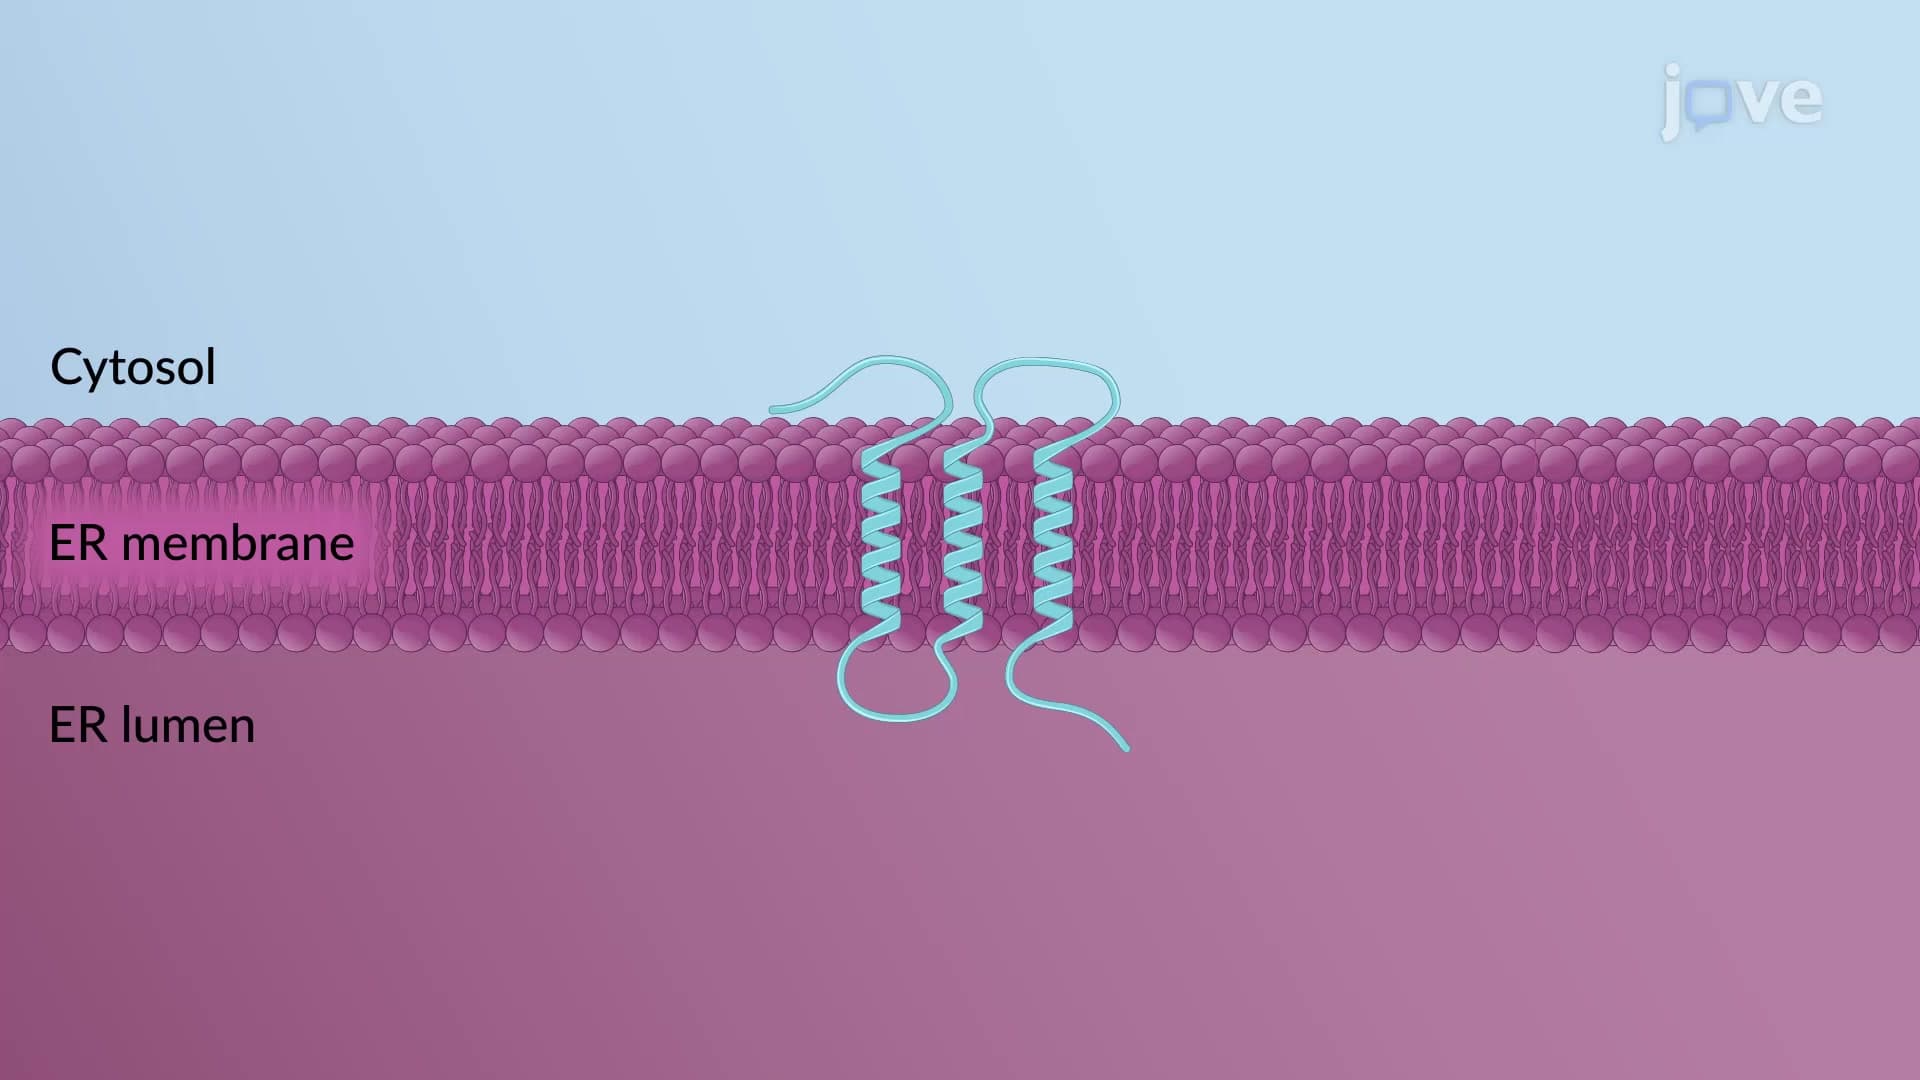
Insertion of Multi-pass Transmembrane Proteins in the RER
The multipass transmembrane proteins are the type IV integral membrane proteins with multiple topogenic sequences determining their spatial arrangement in the ER membrane. Nearly all multipass proteins lack a cleavable signal sequence and use...
 01:45
Tail-anchoring of Proteins in the ER Membrane