Conformational transitions using molecular dynamics with minimum biasing
Related Experiment Video
Updated: Nov 7, 2026
Exploring Caspase Mutations and Post-Translational Modification by Molecular Modeling Approaches
Published on: October 13, 2022
1Department of Biochemistry, University of Alabama, School of Medicine, Birmingham 35294.

Abstract:
The molecular dynamics algorithm (MD), which simulates intramolecular motions on the subnanosecond timescale, has been modified to allow the investigation of slow conformational transitions that do not necessarily occur spontaneously in MD simulations. The method is designated CONTRA MD (CONformational TRAnsitions by Molecular Dynamics with minimum biasing). The method requires the prior definition of a single conformational variable that is required to vary monotonically from an initial conformation to a final target conformation. The simulation is broken up into a series of short free MD segments, and we determine, after each segment of MD, whether or not the system has evolved toward the final conformation. Those segments that do not move the system in that direction are deleted. Those that do move it toward the final conformation are patched together sequentially to generate a single representative trajectory along the transition pathway. The CONTRA MD method is demonstrated first by application to the simultaneous C2'-endo to C3'-endo repucker and anti to syn N-glycosidic torsion transitions in 2'-deoxyadenosine and then to the large-scale bending in phenylalanine transfer RNA.
More Related Videos
Related Concept Videos
 00:52
¹H NMR of Conformationally Flexible Molecules: Temporal Resolution
 01:15
¹H NMR of Conformationally Flexible Molecules: Variable-Temperature NMR
 01:58
Cooperative Allosteric Transitions
01:58Cooperative Allosteric Transitions
01:58Cooperative Allosteric Transitions
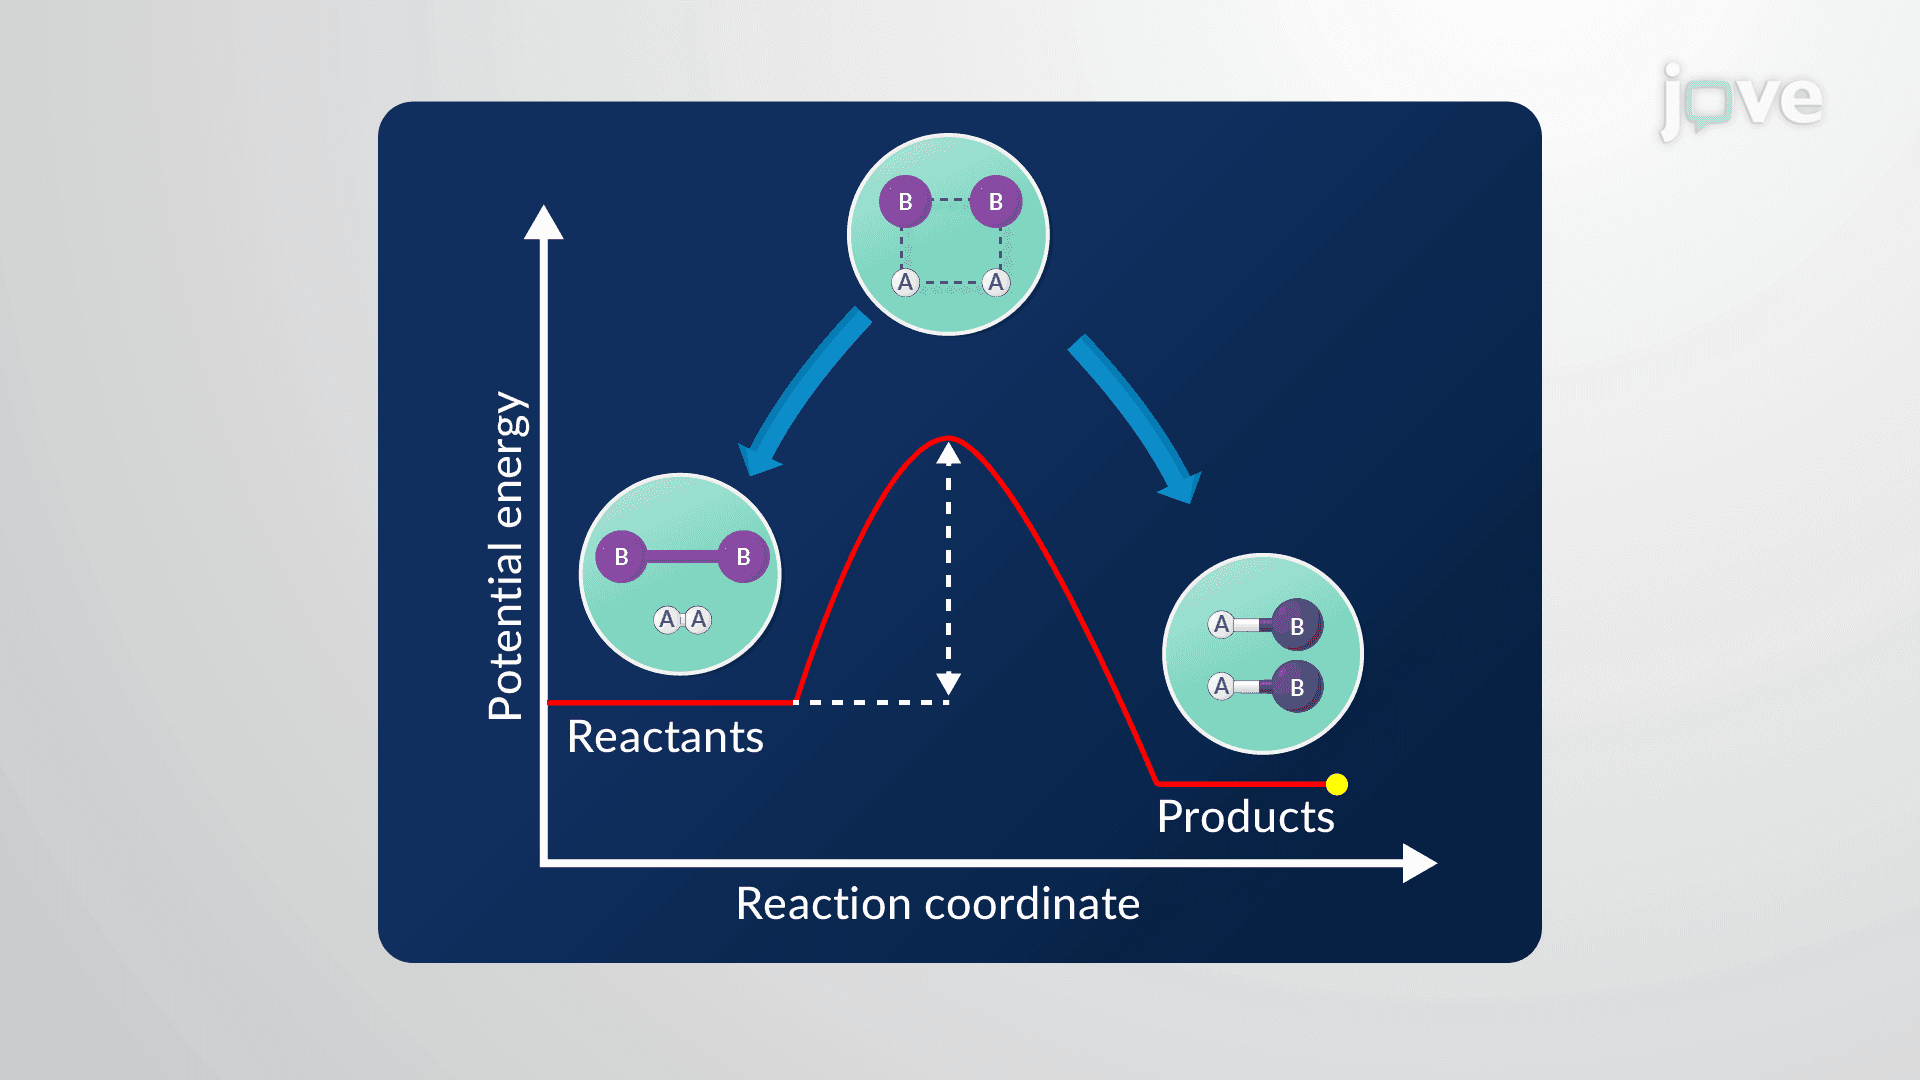 01:25
Transition State Theory